FDA

Why FDA Recalls Cancer-Linked Blood Pressure Medication

US rule body, FDA, said drug firms pull some heart pills back. These pills had bad stuff that might cause bad growths. Lots of folks use these drugs to keep pulse low. This news makes them fret.

Pills That Face Pull?

Pills pulled are a type named ARBs. Docs give these for high pulse and weak heart. They work by making blood paths wide. This lets the heart push blood out with less strain.

Bad pills use names like valsartan, losartan, and irbesartan. Many US folks take these each day. They keep the pulse in check.


What’s the Bad Stuff?

FDA found a dirt called nitrosamine in the pills. Docs say nitrosamines could make men sick with growths if folks use high sums for a long time. These germs can pop up when drugs are made in set ways.

Dirt found were:

  • NDMA
  • NDEA
  • NMBA

How Did This Occur?

The dirt seems to come from shifts in how drugs are made. This is most true in plants far from here. The FDA works with other land rule groups to find the source. They need firms to fix things now.

The rule body said they found the slip in their normal checks. Since finding it, FDA looks at more firms and more drugs.

What Should Sick Folks Do?

FDA says: NO WAY stop pills! Talk to your doc first! This is key for big facts:

  1. Now risk vs. Maybe risk: Bad pulse now can make a heart stop or cause a brain bleed. This risk is fast. The growth risk is slow. The fast risk wins.
  2. Not all pills suck: Lots of heart pills, even in the same group from new firms, are fine.
  3. Other drugs exist: Docs can give new pills or switch to safe lots of the old pills.

Steps Sick Folks Must Take:

  • Keep on with your pill but wait for your doc’s word.
  • Call your care giver to ask if your pill dose is bad.
  • Look at the FDA site for lists of bad pills.
  • Ask your drug shop keeper for news on your script.
  • Do not swap pills or use meds from friends.**

How Big Is the Risk?

FDA says the bad dirt sums are small. The chance for bad growths comes from long use. The rule body still checks what this means for health.

Gurus say the risk for bad growths rests on some facts:

  • How much dirt is in your pill batch
  • How long you took the bad pill
  • Facts on the sick man and his own health

FDA set how much dirt is okay. They press firms to meet these safe sums now.

Firms Act Now

Drug firms work with FDA to do this:

  • Pull back bad pill lots
  • Look deep at how they make stuff
  • Make shifts to stop dirt next time
  • Test both final drugs and raw stuff for bad bits

Both known brands and cheap pills face this pull. Firms work to make sure new pills pass all safe checks.

FDA Keeps Looking

FDA keeps on with this work:

  • Watch the flow of drugs in the land
  • Test drugs for dirt that pops up
  • Tell the world when new pills get pulled
  • Work with rule groups on far land plants
  • Judge the long run health cost for men who took bad pills

What This Means for Care

This shows why some things are vital:

  • Good checks in drug making: Hard tests and watch are key for safe drugs.
  • Watch the whole chain of stuff: Lots of drugs come from far lands. This needs world groups to share safe rules.
  • Talk to sick folks: Care givers must tell folks things right, not cause fright.
  • Rule body must stay sharp: Keep testing and watching to spot bad things fast.

What’s Next

FDA hopes things get good as firms fix ways and the rule body makes checks more tight. But folks might find some heart pills hard to get for a bit. Bad drugs pull out. Safe drugs move in.

The rule body works with firms to keep drugs near, but safe first. In some cases, FDA lets pills with just a bit of dirt stay out if taking them now is way more good than the slight risk they pose.

Wrap Up

Finding dirt that might cause bad growths in heart pills is scary. But sick folks must see that high pulse is a fast, great risk. FDA, docs, and drug makers join up to fix this and keep folks safe.

If you take heart pills, stay up to date. Talk clear with your care giver. Keep taking your pills as told, unless your doc says stop. Your health is the top aim as this mess gets fixed and watched.

Why FDA Recalls Cancer-Linked Blood Pressure Medication Read More »

Are there any FDA-approved OTC Sex-enhancement Pills in USA

Recent developments in sexual health products have created confusion about which sex-enhancement pills or treatments are actually FDA-approved and available without a prescription in the United States. Many consumers searching for accessible solutions to sexual performance issues face a marketplace filled with products making bold claims. This comprehensive guide examines the current landscape of over-the-counter sexual enhancement options, separating FDA-approved treatments from potentially dangerous alternatives.


The Current State of OTC Sex-Enhancement Products

Despite the abundance of products marketed as “sex-enhancement pills” on store shelves and online retailers, the reality is that there are extremely limited FDA-approved over-the-counter options available. Most products claiming to enhance sexual performance that don’t require a prescription fall into the dietary supplement category, which does not undergo the same rigorous FDA approval process as medications.

Eroxon: The First FDA-Cleared OTC Treatment

In June 2023, the FDA cleared MED3000, marketed as Eroxon, as the first over-the-counter treatment for erectile dysfunction. However, it’s important to note that Eroxon is not a pill but a topical gel that is applied directly to the head of the penis.

Eroxon works through a physical mechanism rather than pharmacological action. When applied, it creates an initial cooling sensation followed by warming, which stimulates nerve endings and increases blood flow to facilitate an erection. Clinical trials found that 65% of men who used the product achieved an erection within 10 minutes, significantly faster than traditional prescription ED medications which typically take around 30 minutes to work.

The FDA considers Eroxon a medical device rather than a medication, and it received “FDA cleared” status, a designation typically used for medical devices that have been reviewed for safety and effectiveness.


The Truth About “Sex-Enhancement Pills” Sold as Supplements

Hidden Ingredients and FDA Warnings

Many products marketed as “all-natural” sex-enhancement pills actually contain undisclosed pharmaceutical ingredients. The FDA has issued hundreds of warnings about unregulated amounts of prescription medications found in these supplements.

For example, the FDA has warned consumers about numerous products including:

1. “Boost — Ultra Sexual Enhancement Formula” which laboratory analysis confirmed contains sildenafil, the active ingredient in Viagra.

2. “Rhino 7S Type F3 7000,” marketed for sexual enhancement, which contains sildenafil and acetaminophen, potentially creating serious health risks.

3. “Control Capsules,” another product promoted for sexual enhancement, which also contained undeclared sildenafil.

Health Risks of Unauthorized Sex-Enhancement Pills

These undeclared ingredients pose significant health risks. Sildenafil may interact with nitrates found in some prescription drugs such as nitroglycerin and may lower blood pressure to dangerous levels. This is particularly concerning for men with diabetes, high blood pressure, high cholesterol, or heart disease who often take nitrates.

The FDA considers this a growing trend of products marketed as dietary supplements with hidden drugs and chemicals. These products are typically promoted for sexual enhancement, weight loss, and body building, and are often represented as being “all natural” despite containing potent pharmaceutical ingredients.


Legitimate Options for Sexual Enhancement

Prescription Medications

FDA-approved prescription medications for erectile dysfunction remain the safest and most effective pharmaceutical options for treating sexual performance issues. These include:

  • Sildenafil (Viagra)
  • Tadalafil (Cialis)
  • Vardenafil (Levitra)
  • Avanafil (Stendra)

These medications require consultation with a healthcare provider to ensure they’re appropriate based on individual health conditions and medications.


Eroxon Gel: The OTC Alternative

For those seeking an over-the-counter option, Eroxon gel represents the only FDA-cleared choice available without a prescription. As a non-medicated product, Eroxon has minimal side effects and doesn’t have the same contraindications as oral ED medications. Side effects were minimal in clinical trials, with headaches and burning or irritation of the penis occurring in only 1% to 3% of men.

Eroxon is available for purchase online or at pharmacies without a prescription. It’s designed for men age 22 and older and comes in single-use tubes.


How to Identify Safe Sexual Enhancement Products

When considering any sex-enhancement product, consumers should:

[1] Look for FDA approval or clearance
[2] Consult with a healthcare provider before trying new products
[3] Be wary of products making extraordinary claims or marketed as “all natural”
[4] Check the FDA’s list of tainted sexual enhancement products
[5] Report adverse effects to the FDA’s MedWatch program

Despite the vast market of products claiming to be effective sex-enhancement pills, there are currently no FDA-approved over-the-counter pills for sexual enhancement in the United States. The only FDA-cleared over-the-counter option is Eroxon, which is a topical gel rather than a pill.

Many products marketed as sex-enhancement pills contain undisclosed prescription medications that could pose serious health risks. Consumers should exercise caution, consult healthcare providers, and consider FDA-approved prescription options or the recently cleared Eroxon gel instead of unregulated supplements.

For those experiencing erectile dysfunction or other sexual health concerns, speaking with a healthcare provider remains the safest first step toward finding effective, medically sound treatment options rather than turning to unregulated sex-enhancement pills that may contain hidden and potentially harmful ingredients.

Are there any FDA-approved OTC Sex-enhancement Pills in USA Read More »

How long does it take for a drug to get FDA approval?

The journey from laboratory discovery to pharmacy shelf is a lengthy and complex process for new medications. According to recent data, the average timeline for drug approval ranges from 8.5 to 12 years, with considerable investment of time and resources at each step. The FDA’s rigorous evaluation process ensures that only safe and effective drugs reach American consumers, but this thoroughness comes at the cost of time. This comprehensive article examines how long it actually takes for drugs to receive FDA approval, the factors that influence these timelines, and recent trends that are reshaping the approval landscape.


The Complete Drug Development Journey

The full research, development, and approval process for a new drug typically spans between 12 to 15 years from initial discovery to market availability. This extensive timeline reflects the meticulous nature of pharmaceutical development, where safety and efficacy must be definitively established before a drug can be made available to the public. More recent estimates suggest an average of 8.5 years for a drug to progress from early laboratory discovery to final approval for human use.
This lengthy process comes with substantial financial implications. Studies have found varying costs, with some research placing the average development cost at $2.8 billion, while a 2020 study determined the average cost to be $1.3 billion, with a median cost of $985 million to bring a new drug to market. These figures highlight the significant investment pharmaceutical companies must make before seeing any return, explaining in part why the industry pushes for efficiency in the approval process.

The timeline begins with extensive laboratory research, often building upon preliminary work from academic institutions, government bodies like the National Institutes of Health, and previous development studies. Only after promising results in the laboratory does the process advance to animal testing and eventually human trials.


The Phases of FDA Drug Approval

The FDA drug approval process consists of several distinct phases, each with its own timeline and requirements.


Pre-Clinical Research and IND Application

Before any testing in humans can begin, pharmaceutical companies must conduct extensive laboratory and animal studies to demonstrate that their drug candidate is reasonably safe for human trials. This preclinical phase is crucial for understanding how the drug works and identifying potential toxicity issues.

Once a sponsor has sufficient preclinical data, they must submit an Investigational New Drug (IND) Application to the FDA. This application details the drug’s composition, manufacturing process, results from animal testing, and the planned protocol for human clinical trials. The FDA reviews this application to ensure that participants in clinical trials won’t be exposed to unreasonable risks.


Clinical Research Phase

The clinical research phase involves testing the drug in humans and typically proceeds through three phases:


Phase 1: Usually involves a small number of participants (20-80 people) to evaluate the drug’s safety, determine safe dosage ranges, and identify side effects. This initial phase focuses primarily on safety rather than effectiveness.

Phase 2: Involves a larger group of participants to further assess safety and begin evaluating effectiveness for a specific medical condition.
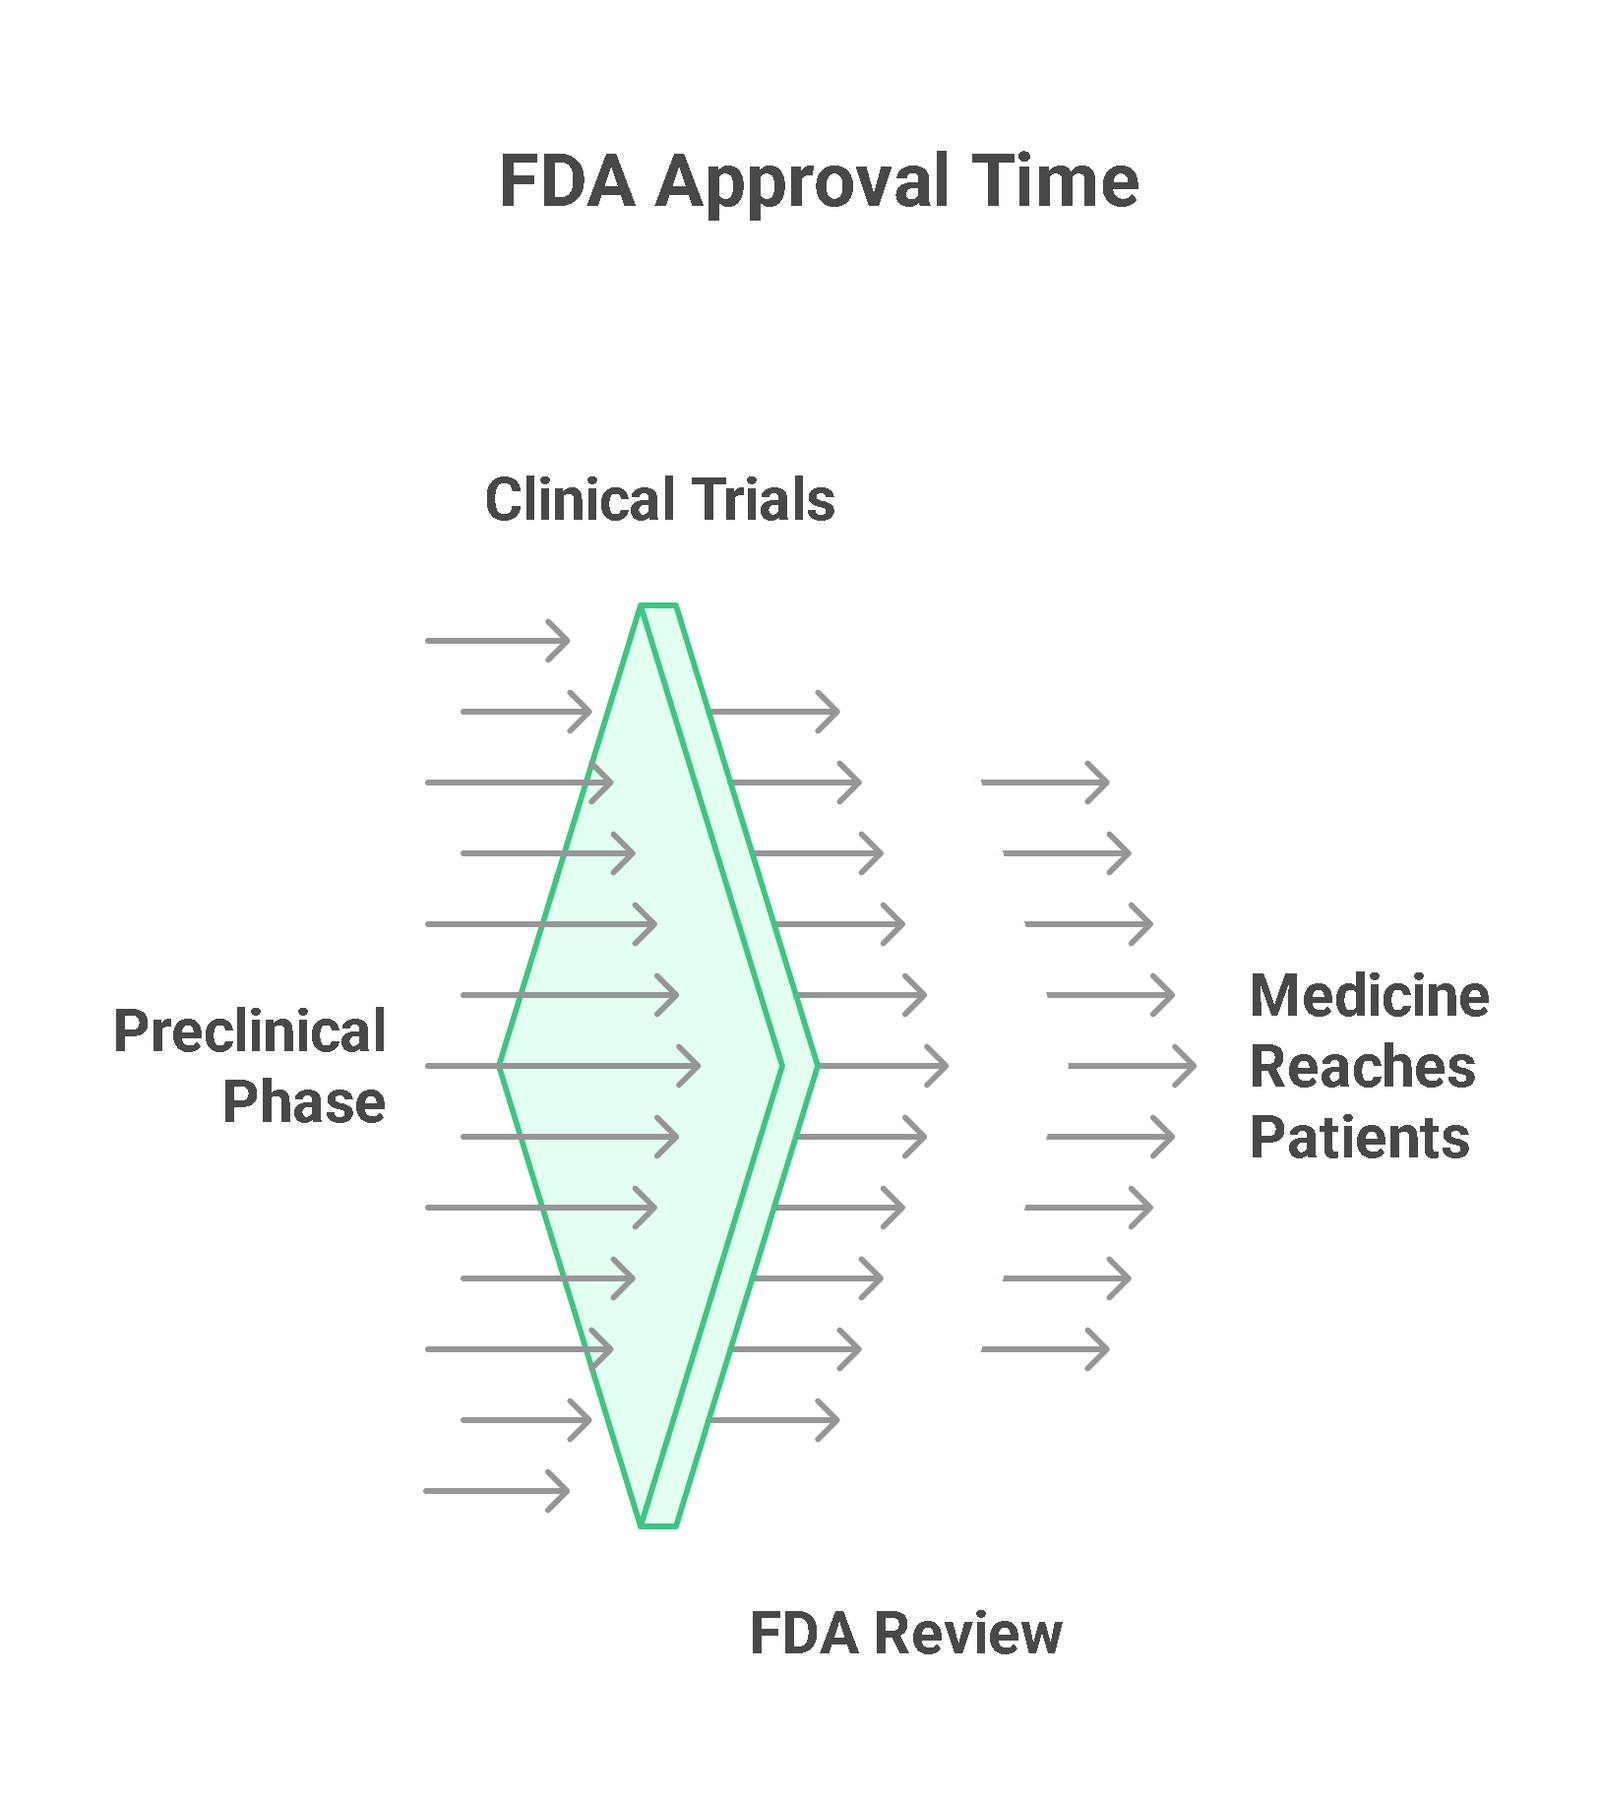
Phase 3: Involves hundreds or thousands of participants to gather more information about safety and effectiveness, monitor side effects, compare the drug to existing treatments, and collect information for appropriate labeling.

These clinical trials represent the longest portion of the drug development process, with the complete clinical development time for a typical innovative drug averaging 9.1 years.


New Drug Application (NDA) Review

After completing clinical trials, the company submits a New Drug Application (NDA) to the FDA. This comprehensive application includes all animal and human test results, manufacturing details, proposed labeling, and other critical information.


A team of FDA scientists, including physicians, statisticians, chemists, pharmacologists, and other experts, then reviews all components of the NDA to determine if the drug’s benefits outweigh its known risks. This review process has two potential timelines:


– Standard Review: The FDA aims to take action within 10 months of receiving the application.
– Priority Review: For drugs that would significantly improve treatment, diagnosis, or prevention of serious conditions, the FDA aims to take action within 6 months.


Post-Marketing Surveillance

Even after approval, the FDA continues to monitor drugs for safety concerns. This ongoing surveillance helps identify adverse reactions that may not have been apparent during clinical trials.


Standard vs. Priority Review: Fast-Tracking Crucial Medications

The FDA has implemented several pathways to expedite the review process for promising medications, particularly those addressing serious conditions with unmet medical needs.

Under standard review, the FDA typically takes 10 months to review a new drug application. However, under priority review, this timeline is shortened to approximately 6 months. This accelerated timeline is reserved for medications that offer significant improvements in safety or effectiveness compared to current treatments.

All six major regulatory agencies worldwide now offer expedited processes to hasten the review of promising new drugs. The FDA’s extensive use of these facilitated regulatory pathways (FRPs) contributes to its position as the agency with the shortest median approval time (244 days) among major regulatory authorities.

In exceptional cases, approval can be remarkably swift. For instance, blinatumomab and remdesivir received FDA approval in just 75 and 76 days, respectively. However, it’s worth noting that remdesivir benefited from a prior emergency use authorization, and blinatumomab initially received accelerated approval that was later converted to full approval.


Recent Trends in FDA Approval Times

In recent years, there has been a clear trend toward faster FDA reviews. According to research published in 2020, the median review time for standard drug applications in 2018 was 10.1 months, significantly shorter than the 2.8 years it took for standard and priority applications in earlier periods.


Comparatively, the FDA has the shortest median approval time (244 days) among six major regulatory authorities, followed by Health Canada (306 days), Japan’s PMDA (313 days), Australia’s TGA (315 days), Europe’s EMA (426 days), and Swissmedic (470 days). This difference likely stems from the FDA’s extensive use of facilitated regulatory pathways designed to expedite promising treatments.

However, some researchers have raised concerns that faster approvals may be associated with reliance on weaker evidence. This highlights the delicate balance the FDA must maintain between speed and thoroughness in its evaluation process.


Case Studies in Rapid Approval

Some drugs have received extraordinarily fast approvals:

– Blinatumomab: Approved in just 75 days, though it initially received accelerated approval that was later converted to full approval.
– Remdesivir for COVID-19: Approved in 76 days after submission, benefiting from a prior emergency use authorization.
– Cabazitaxel: Approved 78 days after completion of a rolling NDA submission.

These rapid approvals typically involve therapies for serious conditions with significant unmet needs, and often utilize special regulatory provisions such as rolling reviews, where the FDA reviews portions of an application as they become available rather than waiting for the complete submission.


Variations by Drug Type and Therapeutic Area

Approval timelines can vary significantly depending on the type of drug and its therapeutic area.

Recent data shows that antiviral products had notably shorter development times compared to other product classes. This may reflect both the urgent public health need for certain antivirals and the relatively straightforward endpoints used to measure their efficacy.

The top five therapeutic areas by number of approvals across major regulatory agencies between 2016-2020 were:

1. Anti-cancer and immunomodulators
2. Anti-infective therapies
3. Blood and blood-forming organs
4. Nervous system treatments
5. Alimentary and metabolism drugs


Among these categories, anti-infective therapies were approved marginally faster with an overall median of 296 days, compared with 340 days for anti-cancer and immunomodulators. However, there is significant variation in development times within each therapeutic class, suggesting that factors beyond the disease category influence approval timelines.

Orphan drugs, which treat rare diseases, often follow different approval pathways. In Japan, all orphan drugs approved went through expedited review due to incentives to address unmet needs, resulting in a rapid median approval time of 200 days. Similarly, in the United States, 81% of orphan products were approved through expedited review, with a median approval time of 234 days.

The journey from laboratory discovery to FDA approval is lengthy and complex, typically taking between 8.5 to 12 years. While the standard review process takes about 10 months from NDA submission, the entire development timeline—including discovery, preclinical research, and clinical trials—stretches much longer.


Recent trends show that the FDA has been approving drugs faster than ever before, with a median approval time of 244 days—the shortest among major regulatory authorities. Priority review, which is granted to drugs that offer significant therapeutic advances, further shortens this timeline to approximately 6 months.


However, approval times vary considerably based on therapeutic area, with antivirals and certain orphan drugs often receiving faster approvals than other categories. The FDA’s implementation of various expedited pathways has contributed significantly to these shorter timelines, particularly for medications addressing serious unmet medical needs.


As the FDA continues to balance thoroughness with efficiency, the drug approval process remains a critical safeguard ensuring that American consumers have access to safe and effective medications, even as the agency works to make promising new treatments available as quickly as possible.

How long does it take for a drug to get FDA approval? Read More »

What are the latest FDA regulations for drug manufacturing?

The U.S. Food and Drug Administration has recently introduced significant updates to drug manufacturing regulations, emphasizing advanced technologies, batch uniformity, and quality assurance. In early 2025, the FDA released new guidance documents that reflect the agency’s evolving approach to pharmaceutical production oversight while maintaining its foundational commitment to public health protection through rigorous quality standards. These regulatory developments represent important considerations for pharmaceutical manufacturers navigating compliance requirements in an increasingly innovative industry landscape.


Current Good Manufacturing Practice Regulations

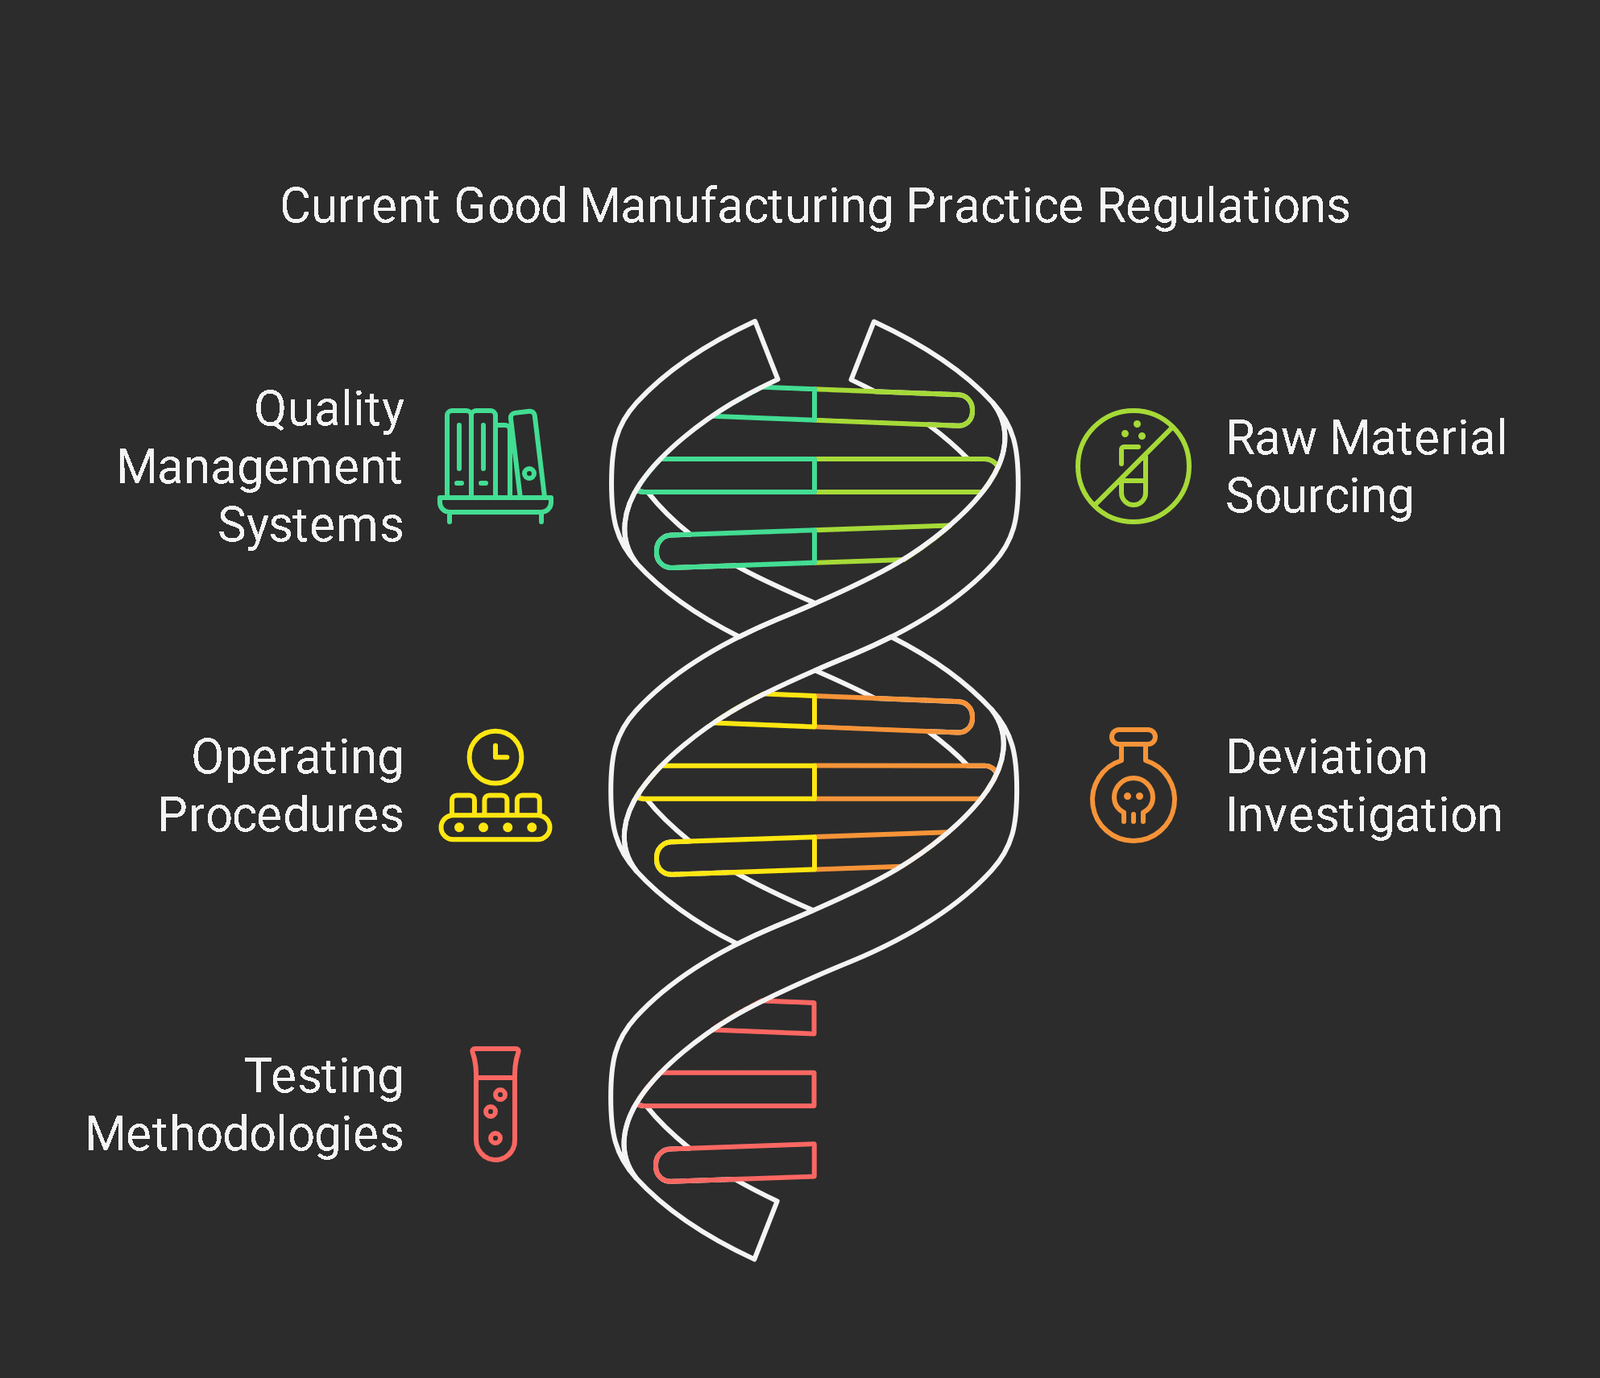
Current Good Manufacturing Practice (CGMP) regulations constitute the cornerstone of pharmaceutical quality assurance in the United States. These regulations, enforced by the FDA, establish minimum standards for manufacturing, processing, packing, and holding human and animal drug products to ensure their safety, efficacy, and quality. CGMP provides comprehensive frameworks for quality management systems, appropriate raw material sourcing, robust operating procedures, quality deviation investigation protocols, and reliable testing methodologies. The regulations were deliberately designed with flexibility to accommodate technological advancements while maintaining stringent quality standards, allowing pharmaceutical companies to implement scientifically sound design, processing methods, and testing procedures appropriate to their specific operations.

The adaptability of CGMP regulations has become increasingly important as the pharmaceutical industry embraces technological innovation. These foundational requirements, codified in 21 C.F.R. Parts 210 and 211, were first issued in 1978 but continue to serve as the primary regulatory mechanism for ensuring pharmaceutical quality despite significant technological evolution since their inception. Compliance with these standards remains mandatory, as failure to meet CGMP requirements may result in a drug being classified as adulterated under Section 501(a)(2)(B) of the Federal Food, Drug, and Cosmetic Act. This classification carries significant legal and business implications for pharmaceutical manufacturers, underscoring the critical importance of understanding and implementing current regulatory expectations.


Recent Interpretive Guidance for CGMP Compliance

On January 6, 2025, the FDA released a draft guidance document titled “Consideration for Complying with 21 C.F.R. 211.110” that provides clarity on the agency’s interpretation of general requirements for drug product manufacturing. This guidance document specifically addresses 21 C.F.R. § 211.110, which requires manufacturers to conduct in-process controls, tests, and examinations to prevent contamination and monitor quality attribute changes in in-process materials. The draft guidance elaborates on FDA’s expectations regarding these requirements and introduces considerations specific to advanced manufacturing technologies, demonstrating the agency’s commitment to supporting innovation while maintaining rigorous quality standards.


The draft guidance emphasizes that manufacturers should develop scientific, risk-based strategies for sampling and testing at critical manufacturing process points to ensure consistent product quality throughout production. It encourages the implementation of real-time quality monitoring, process analytical technologies (PAT), and continuous manufacturing systems to enhance operational efficiency and product quality. Importantly, the guidance recommends that process models be paired with in-process testing or examination to ensure compliance with regulatory requirements, reflecting a balanced approach to technological innovation and traditional quality assurance methodologies.


Advanced Manufacturing Technologies Designation Program

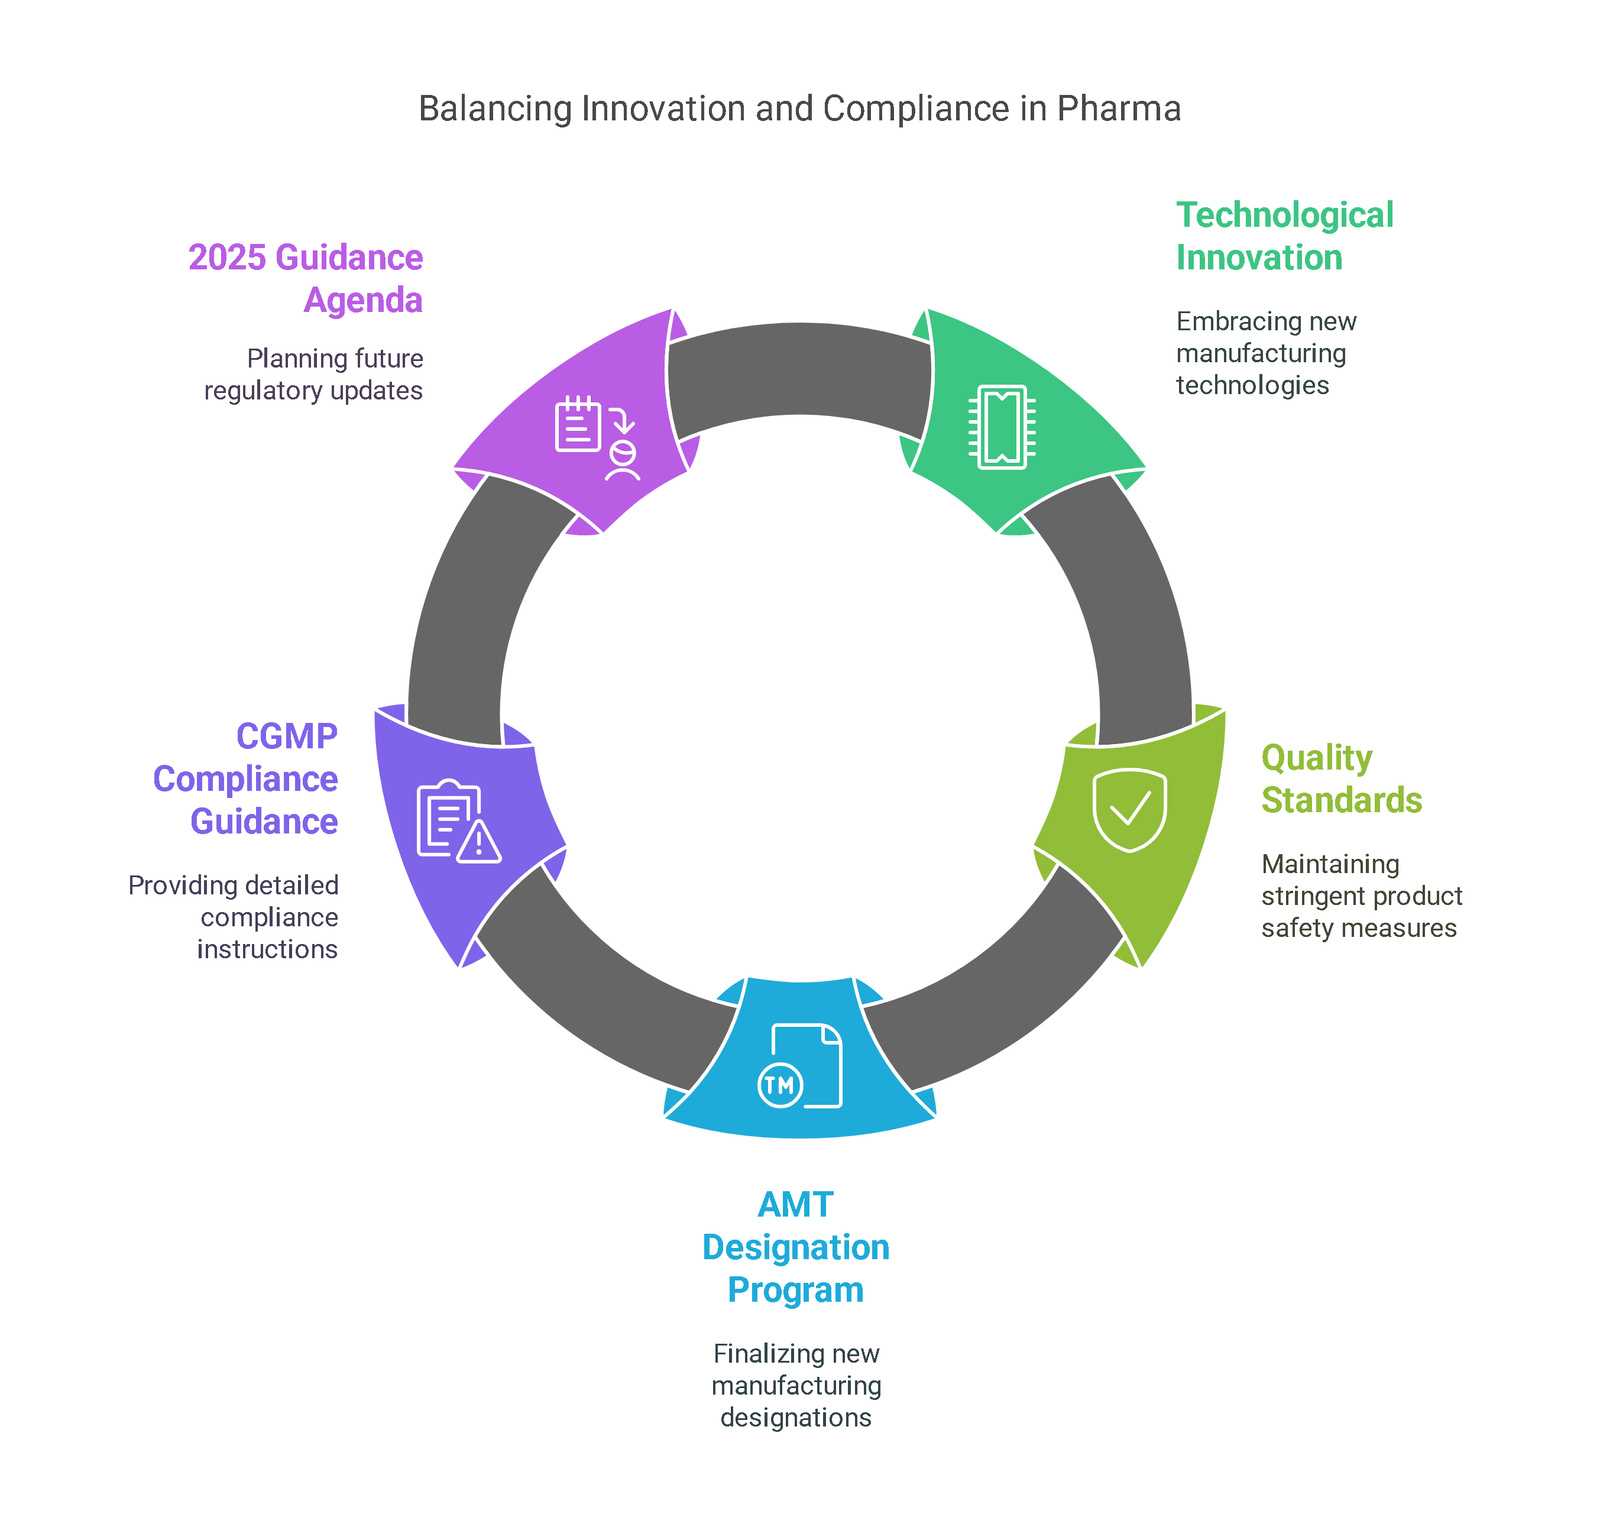
In a significant development for pharmaceutical manufacturing innovation, the FDA published the final guidance for its Advanced Manufacturing Technologies (AMT) Designation Program on January 2, 2025. This program, mandated under section 506L of the Federal Food, Drug, and Cosmetic Act, aims to facilitate the development of drugs manufactured using designated advanced manufacturing technologies. The finalized guidance provides comprehensive recommendations for organizations interested in participating in this program, which was initially proposed in draft form on December 13, 2023.


The FDA defines “advanced manufacturing” as “an innovative pharmaceutical manufacturing approach or technology that has the potential to improve the reliability and robustness of the manufacturing process and supply chain and increase timely access to quality medicines”. This broad definition encompasses novel technological approaches, innovative applications of established techniques, and the implementation of production methods in new domains where defined best practices or experience may not yet exist. Through the AMT Designation Program, the FDA aims to encourage early adoption of technologies that can enhance manufacturing process reliability and potentially reduce drug development timelines while increasing the supply of critical medications.


 Benefits and Implementation of the AMT Program

The AMT Designation Program offers several significant benefits to pharmaceutical manufacturers adopting innovative production technologies. Program participants gain access to early-stage discussions with FDA experts, allowing potential regulatory approval challenges to be addressed proactively. Additionally, the program provides for expedited assessments of applications involving AMT-designated technologies, potentially streamlining the path to market for drugs manufactured using these innovative methods. Perhaps most importantly, the integration of advanced control strategies through this program helps manufacturers meet demand for essential medicines while minimizing quality risks, supporting both industry efficiency and public health objectives.


The detailed guidance document outlines eligibility criteria for AMT designation and provides a structured framework for the submission and assessment process for designation requests. Notably, the guidance acknowledges that technology developers who are not drug application holders may seek AMT designation by providing data generated with model drugs to demonstrate the technology’s parameters, limitations, and intended context of use. This provision encourages technology innovation across the pharmaceutical ecosystem, potentially broadening the impact of the program beyond established drug manufacturers to include specialized technology developers and other industry stakeholders.


Upcoming Regulatory Developments: The 2025 Guidance Agenda

The FDA’s Center for Drug Evaluation and Research (CDER) has published an ambitious guidance agenda for calendar year 2025, indicating the agency’s regulatory priorities across numerous pharmaceutical manufacturing and quality categories[2]. This agenda includes planned guidance documents addressing various aspects of pharmaceutical development, manufacturing, and quality control, providing valuable insight into the evolving regulatory landscape for drug manufacturers.


Several forthcoming guidance documents directly relate to manufacturing practices and quality assurance, including guidance on “Laboratory Testing of Drugs Held in Interstate Commerce: Compliance with CGMP” and a revised draft guidance on “PET Drugs – Current Good Manufacturing Practice (CGMP)”. Additional planned guidance documents address areas such as stability recommendations for manufacturing facilities, guidelines for establishing impurity limits for antibiotics, and current good manufacturing practice for medical gases. The comprehensive nature of this guidance agenda reflects the FDA’s ongoing commitment to providing clarity and direction across the full spectrum of pharmaceutical manufacturing activities.


The guidance agenda also includes numerous documents focused on specialized manufacturing considerations for biological products and generic drugs. These include guidance on “Biosimilar and Interchangeable Biosimilar Products: Considerations for Container Closure Systems and Device Constituent Parts” and various documents addressing bioequivalence studies and testing methodologies for abbreviated new drug applications. This specialized guidance demonstrates the FDA’s recognition of unique manufacturing challenges across different pharmaceutical product categories and its commitment to providing tailored regulatory frameworks appropriate to diverse manufacturing contexts.


Implications for Pharmaceutical Manufacturers

The recent and planned regulatory developments have significant implications for pharmaceutical manufacturers. The emphasis on advanced manufacturing technologies and process models demonstrates the FDA’s recognition of technological evolution in pharmaceutical production while maintaining its fundamental commitment to product quality and safety. Manufacturers are encouraged to embrace innovation but must ensure that new technologies are implemented within compliant frameworks that satisfy established quality requirements.


The draft guidance on complying with 21 CFR 211.110 provides valuable clarity on the agency’s expectations regarding batch uniformity and in-process controls. Manufacturers should review their existing sampling and testing protocols against this guidance to ensure alignment with current regulatory thinking. Particular attention should be given to the integration of process models with traditional testing methodologies, as the guidance specifically recommends this balanced approach to quality assurance in advanced manufacturing contexts.


For manufacturers interested in implementing novel production technologies, the finalized AMT Designation Program offers a structured pathway for engagement with the FDA. The program’s benefits, including early regulatory interaction and expedited application assessment, may provide significant advantages for organizations at the forefront of manufacturing innovation. However, participation requires thorough preparation and documentation to demonstrate that proposed technologies meet the program’s eligibility criteria and possess sufficient maturity for consistent and reliable manufacturing in their intended contexts of use.


The Future of Pharmaceutical Manufacturing Regulation

The FDA’s recent regulatory updates reflect a carefully balanced approach to pharmaceutical manufacturing oversight that embraces technological innovation while maintaining stringent quality standards. The finalization of the AMT Designation Program and the release of interpretive guidance for CGMP compliance demonstrate the agency’s commitment to adapting regulatory frameworks to contemporary manufacturing realities while preserving the fundamental quality assurance principles that protect public health.

As pharmaceutical manufacturing continues to evolve, the FDA’s regulatory approach is likely to develop in parallel, with additional guidance documents providing clarity on emerging technologies and methodologies. The comprehensive 2025 guidance agenda indicates the breadth of regulatory considerations the agency plans to address in the coming year, providing a roadmap for manufacturers seeking to anticipate compliance requirements. By maintaining awareness of these regulatory developments and proactively adapting manufacturing practices to align with current FDA expectations, pharmaceutical manufacturers can navigate the complex regulatory landscape while capitalizing on opportunities for innovation and efficiency in drug production.


The pharmaceutical industry stands at an inflection point where advanced technologies offer unprecedented opportunities for manufacturing improvement while regulatory frameworks continue to emphasize the paramount importance of product quality and safety. Successful manufacturers will be those that effectively balance innovation with compliance, leveraging new technologies within well-designed quality systems that satisfy both current requirements and emerging regulatory expectations for pharmaceutical production excellence.

What are the latest FDA regulations for drug manufacturing? Read More »