Quality Management

Essential GMP Principles for Pharma Industry Professionals

Dr. Michael Lee

CA, USA


Introduction

Medicines are unlike most products people buy. Patients cannot see, taste, or test a tablet or injection to know whether it is safe and effective. They trust that the product contains the right ingredient, in the right amount, free from harmful contaminants, every time. Good Manufacturing Practice (GMP) is the system that earns and protects that trust.

GMP is the part of quality assurance that ensures medicinal products are consistently produced and controlled to the quality standards appropriate for their intended use. It is not a single procedure or a checklist for inspectors. It is a way of working that applies to everyone, from the operator weighing raw materials to the senior manager approving a budget.

This article outlines the essential GMP principles every pharmaceutical professional should understand, why each matters, and how to apply them in daily work.


1. Why GMP Exists

GMP regulations grew out of tragedies. Contaminated and poorly made medicines have harmed and killed patients throughout the twentieth century, and each major incident pushed regulators to tighten the rules. The Elixir Sulfanilamide poisoning in the United States in 1937, the thalidomide disaster of the early 1960s, and later contamination events involving sterile products and diethylene glycol in liquid medicines all shaped modern requirements.

The lesson from this history is that quality cannot be tested into a product. Final testing examines only a small sample and can miss problems that are unevenly distributed in a batch. Quality must be built in through controlled materials, validated processes, qualified equipment, trained people, and reliable records. GMP provides the framework for doing exactly that.


2. The Foundation: Pharmaceutical Quality System and Quality Management

GMP operates inside a broader Pharmaceutical Quality System (PQS), described in ICH Q10. The PQS defines how the organization manages quality: through leadership commitment, clear responsibilities, documented procedures, and continual improvement.

Core expectations include:

  • Management responsibility. Senior leaders must provide the resources, structure, and support for compliance, and they remain accountable for it.
  • An independent quality unit. The quality function must have the authority to approve or reject materials, batches, and procedures without pressure from production or commercial teams.
  • Risk-based decision-making. Resources and controls are directed to where the risk to patients is highest.
  • Continual improvement. Deviations, complaints, audit findings, and trends feed back into better processes.

Without this foundation, individual GMP practices tend to become isolated rules followed inconsistently.


3. Personnel: Training, Hygiene, and Responsibility

People are the most important and the most variable element in any manufacturing process. GMP therefore sets clear expectations for them.

Qualified and Trained Staff

Every facility needs enough qualified personnel to perform the work correctly. Employees should have defined roles, job descriptions, and documented training that covers:

  • Initial GMP training on joining the company.
  • Job-specific training on the procedures they perform.
  • Ongoing refresher training, including updates when procedures or regulations change.
  • Specialized training for higher-risk areas such as sterile manufacturing, potent compounds, or laboratory operations.

Training is only effective when it is assessed. Signing an attendance sheet proves presence, not competence. Observation, practical demonstration, and performance review are stronger evidence.

Personal Hygiene and Health

Operators are a potential source of contamination. GMP requires hygiene programs that cover handwashing, appropriate clothing and gowning, restrictions on jewelry and cosmetics in production areas, and prohibitions on eating, drinking, and smoking in manufacturing zones. Employees should report illnesses or open wounds that could compromise product quality. In sterile areas, gowning and aseptic technique are qualified and monitored.

Shared Responsibility

Although the quality unit has formal authority, quality is everyone’s responsibility. Operators should feel empowered to stop a process when something looks wrong, and supervisors should support that decision.


4. Premises and Facility Design

Facilities must be designed, located, constructed, and maintained to suit the operations performed in them. Good design prevents mix-ups, cross-contamination, and errors before they occur.

Key principles include:

  • Logical flow. Materials, personnel, and waste should move in a way that minimizes the risk of contamination and mix-ups.
  • Segregation. Areas for receiving, quarantine, sampling, production, packaging, storage, and quality control should be separated or controlled appropriately.
  • Surfaces and finishes. Walls, floors, and ceilings should be smooth, impervious, and easy to clean and disinfect.
  • Environmental control. HVAC systems regulate temperature, humidity, air quality, and pressure differentials between rooms.
  • Dedicated facilities where needed. Highly sensitizing or hazardous products, such as certain penicillins, cephalosporins, or potent compounds, may require dedicated or strictly segregated facilities.
  • Pest control and maintenance. Buildings should be kept in good repair, with effective pest control programs.

Cleanrooms are classified by airborne particle limits and microbial limits, and sterile operations rely on unidirectional airflow, pressure cascades, and environmental monitoring to maintain control.


5. Equipment: Qualification, Calibration, and Maintenance

Equipment must be suitable for its purpose and must not adversely affect product quality. GMP expects a lifecycle approach:

  • Design and selection. Equipment should be made of materials that do not react with, add to, or absorb from the product.
  • Qualification. Installation Qualification (IQ), Operational Qualification (OQ), and Performance Qualification (PQ) demonstrate that equipment is installed correctly and performs as intended.
  • Calibration. Measuring and control instruments, such as balances, thermometers, and pressure gauges, are calibrated against traceable standards at defined intervals.
  • Preventive maintenance. Scheduled maintenance reduces breakdowns and drift, and records show what was done and when.
  • Status labeling. Equipment should be clearly labeled as clean, in use, under maintenance, or out of service.
  • Cleaning procedures. Written, validated procedures describe how equipment is cleaned between batches and between products.

Computerized systems that control equipment or generate GMP data must also be validated, following the principles of GAMP 5 and Annex 11 of the EU GMP guidelines.


6. Materials Management

Product quality starts with the quality of materials. GMP requires control from the moment a material arrives until it is used or disposed of.

  • Approved suppliers. Suppliers of active substances, excipients, and primary packaging are qualified, and quality agreements define responsibilities.
  • Receipt and quarantine. Incoming materials are inspected, identified, and held in quarantine until released.
  • Sampling and testing. Samples are tested against approved specifications. Identity testing of every container, or a justified alternative approach, guards against mix-ups and adulteration.
  • Storage. Materials are stored under labeled, controlled conditions, with clear separation of quarantined, approved, and rejected stock.
  • Stock rotation. First-expired, first-out (FEFO) or first-in, first-out (FIFO) systems reduce the use of old stock.
  • Traceability. Each container is labeled with its name, batch number, status, and expiry or retest date.
  • Rejected materials. Rejected items are clearly identified and segregated to prevent accidental use.

Because supply chains are global and complex, risk-based supplier oversight, including audits, is increasingly important.


7. Documentation: The Backbone of GMP

An old GMP saying holds that “if it isn’t written down, it didn’t happen.” Documentation describes what to do, and records prove that it was done.

Types of Documents

  • Specifications for materials, intermediates, and finished products.
  • Master formulae and manufacturing instructions that define how a product is made.
  • Standard Operating Procedures (SOPs) that describe routine tasks.
  • Batch manufacturing and packaging records that capture what actually happened for each batch.
  • Logbooks and test records for equipment use, cleaning, calibration, and laboratory work.
  • Validation protocols and reports.

Good Documentation Practices

Documents should be clear, approved, version-controlled, and available at the point of use. Entries should be made at the time of the activity, in indelible ink or validated electronic systems. Corrections should never obscure the original entry: a single line, the date, the initials, and the reason should be recorded. Blank spaces should not be left where data might be added later.

Data Integrity

Regulators place heavy emphasis on data integrity, summarized by the ALCOA+ principles. Data must be Attributable, Legible, Contemporaneous, Original, and Accurate, as well as Complete, Consistent, Enduring, and Available. Electronic systems need unique user accounts, access controls, secure audit trails, and regular audit trail review. Practices such as testing into compliance, deleting raw data, sharing passwords, or backdating records are serious violations and have led to some of the most severe regulatory actions in recent years.


8. Production Controls

Production is where materials become medicines, and GMP requires that it be carried out according to approved, validated instructions.

Essential controls include:

  • Line clearance. Before starting a new batch or product, the area and equipment are checked to confirm that all materials, documents, and labels from the previous operation have been removed.
  • Verified dispensing. Weighing and dispensing are checked by a second person or by a validated electronic system.
  • Defined process parameters. Critical parameters such as mixing time, temperature, compression force, and fill volume are specified, monitored, and recorded.
  • In-process controls. Checks during production, such as tablet weight, hardness, pH, or fill volume, detect problems early.
  • Yield reconciliation. Actual yields are compared with theoretical yields, and unexplained discrepancies are investigated.
  • Prevention of cross-contamination. This involves technical measures such as dedicated equipment, closed systems, and air handling, and organizational measures such as campaign production, cleaning verification, and cleaning validation.
  • Labeling of intermediates and bulk products. Clear identification prevents mix-ups.

Any deviation from approved instructions must be documented and, where appropriate, investigated and approved by the quality unit.


9. Validation and Qualification

Validation provides documented evidence that a process, procedure, or system consistently leads to the expected results.

  • Process validation follows a lifecycle approach: process design, process qualification, and continued process verification. It confirms that the commercial process reliably produces product that meets its specifications.
  • Cleaning validation demonstrates that cleaning procedures remove product residues, cleaning agents, and microbial contaminants to acceptable limits, with limits increasingly derived from toxicological, health-based assessments.
  • Analytical method validation confirms that test methods are accurate, precise, specific, linear, and robust for their intended purpose.
  • Computerized system validation ensures that software is fit for purpose and protects data integrity.
  • Equipment and facility qualification confirms that utilities, equipment, and cleanrooms perform as required.

Validation is not a one-time event. Changes must be evaluated through change control, and revalidation is performed when they could affect product quality.


10. Quality Control and Laboratory Practices

The quality control laboratory verifies that materials and products meet their specifications. GMP expects laboratories to operate with the same discipline as production.

  • Approved methods. Only validated or verified methods are used.
  • Qualified instruments. Equipment is qualified, calibrated, and maintained.
  • Reference standards and reagents. These are properly sourced, labeled, stored, and used within their expiry.
  • Representative sampling. Sampling plans and techniques ensure samples reflect the whole batch or lot.
  • Stability testing. Stability programs establish shelf life and storage conditions and continue after commercial launch.
  • Out-of-specification (OOS) investigations. A failing result must trigger a structured, documented investigation rather than repeated testing until a passing result appears. Invalidating a result requires an assignable, scientifically justified cause.
  • Retention samples. Samples of starting materials and finished products are kept for defined periods to support investigations.

The laboratory must also maintain its own data integrity controls, including audit trail review and restricted access to results.


11. Deviations, CAPA, and Change Control

Even well-run facilities face unexpected events. GMP does not assume perfection. It requires that problems are detected, reported, investigated, and prevented from recurring.

Deviation Management

Any departure from approved procedures or specifications is recorded, assessed for impact on product quality, and investigated in proportion to its risk. Prompt reporting matters. A culture that punishes honest reporting produces hidden problems.

Root Cause Analysis and CAPA

Investigations should identify the true root cause rather than stopping at “operator error.” Tools such as the 5 Whys and fishbone diagrams help. Corrective and Preventive Actions (CAPA) then address the cause, with assigned owners, due dates, and follow-up to confirm that the actions worked.

Change Control

Changes to processes, materials, equipment, facilities, methods, or software are proposed, assessed for risk and regulatory impact, approved by the quality unit, implemented in a controlled manner, and verified afterward. Uncontrolled change is among the most common causes of unexpected quality failures.


12. Packaging and Labeling Controls

Packaging and labeling errors are a leading cause of product recalls. A wrong label or leaflet can cause serious harm, so GMP sets strict controls.

  • Segregation and line clearance between packaging runs.
  • Reconciliation of printed materials, including labels and cartons, with documented investigation of any discrepancy.
  • Verification of printed details, such as batch number, expiry date, and product name, at start-up and at intervals during the run.
  • Automated inspection, such as vision systems and barcode verification, where appropriate.
  • Secure storage and control of printed packaging materials, with destruction of surplus coded items.
  • Serialization and track-and-trace requirements, where applicable, to help protect the supply chain from counterfeit products.

13. Complaints, Recalls, and Product Quality Review

GMP does not end when a batch is released.

  • Complaint handling. Complaints are recorded, evaluated, and investigated, and trends are reviewed. Complaints related to potential quality defects may trigger batch investigations and, where needed, recalls.
  • Recall procedures. Companies must have a tested, documented plan to remove defective products from the market quickly, with clear responsibilities and communication channels.
  • Product Quality Review (PQR) or Annual Product Review (APR). A periodic review of each product examines data such as batch results, deviations, changes, stability, complaints, and trends to confirm the process remains in control and to identify improvements.

These activities close the loop by using real-world performance to improve future manufacturing.


14. Contract Manufacturing and Outsourced Activities

Many companies use contract manufacturers and laboratories. GMP is clear that the contract giver remains responsible for the quality of products made on its behalf.

Good practice includes:

  • Assessing and approving contract acceptors before use.
  • Establishing a written quality agreement that defines responsibilities for manufacturing, testing, change notification, deviations, and release.
  • Conducting periodic audits and performance reviews.
  • Keeping open communication channels for changes and problems.

Outsourcing the activity does not outsource the accountability.


15. Self-Inspection and Audits

Regular self-inspections (internal audits) let a company identify and correct weaknesses before regulators find them. A strong program is risk-based, covers all GMP areas, uses trained and objective auditors, and leads to documented corrective actions with management oversight. Supplier audits and third-party audits complement internal programs.

Effective audits look at how work is actually performed, not just whether procedures exist. Observing operations, speaking with staff, and examining records provide a realistic picture of compliance.


16. Building a Culture of GMP Compliance

Regulatory inspectors increasingly look beyond documents to assess quality culture, meaning the shared attitudes and behaviors toward quality and compliance. A strong culture has recognizable features:

  • Visible leadership commitment, including willingness to delay a shipment rather than compromise quality.
  • Openness. Employees can raise concerns and report mistakes without fear of unfair blame.
  • Accountability. People understand their roles and take ownership of quality.
  • Learning orientation. Errors are studied to improve systems.
  • Consistency. Procedures are followed whether or not an inspector is present.

The practical test of culture is what happens when quality conflicts with schedule or cost. The answer shows what the organization truly values.


17. Everyday Practices for Pharma Professionals

GMP principles become real through daily habits. Some practical guidelines for anyone working in the industry:

  1. Follow the current approved procedure. If it does not work, raise a change request rather than improvising.
  2. Document as you go. Record data at the time of the activity, accurately and completely.
  3. Never falsify or backdate. Honest documentation of a mistake is always better than concealment.
  4. Report deviations promptly. Early reporting allows better containment.
  5. Keep your area clean and organized. Good housekeeping prevents contamination and mix-ups.
  6. Stay current with training. Understand the reasons behind procedures, not only the steps.
  7. Question what seems wrong. If a result, instruction, or situation looks unusual, stop and ask.
  8. Think of the patient. Every action, however small, ultimately affects someone relying on the product.

Conclusion

GMP is often seen as a burden of rules, but its purpose is simple and meaningful: protect patients by ensuring that every medicine is safe, effective, and of consistent quality. The essential principles covered here, including a strong quality system, trained personnel, suitable facilities and equipment, controlled materials, reliable documentation, validated processes, rigorous laboratory practices, effective handling of deviations and change, and a genuine culture of quality, work together as a single system. Weakness in one area can undermine the others.

For pharmaceutical professionals, understanding GMP means understanding not only what the rules require but why they exist. That understanding turns compliance from a checklist exercise into a professional commitment. Regulations will continue to evolve with new technologies such as continuous manufacturing, digital systems, and advanced therapies, but the underlying principles of control, documentation, risk management, and patient focus will remain constant.


Guideline References

  1. ICH Q7 – Good Manufacturing Practice Guide for Active Pharmaceutical Ingredients.
  2. ICH Q8 (R2) – Pharmaceutical Development.
  3. ICH Q9 (R1) – Quality Risk Management.
  4. ICH Q10 – Pharmaceutical Quality System.
  5. US FDA, 21 CFR Part 210 – Current Good Manufacturing Practice in Manufacturing, Processing, Packing, or Holding of Drugs: General.
  6. US FDA, 21 CFR Part 211 – Current Good Manufacturing Practice for Finished Pharmaceuticals.
  7. US FDA (2011) – Guidance for Industry: Process Validation: General Principles and Practices.
  8. US FDA (2018) – Guidance for Industry: Data Integrity and Compliance With Drug CGMP: Questions and Answers.
  9. US FDA (2022) – Guidance for Industry: Investigating Out-of-Specification (OOS) Test Results for Pharmaceutical Production.
  10. European Commission, EudraLex Volume 4 – EU Guidelines for Good Manufacturing Practice for Medicinal Products for Human and Veterinary Use, including Part I (Basic Requirements for Medicinal Products), Annex 1 (Manufacture of Sterile Medicinal Products), Annex 11 (Computerised Systems), and Annex 15 (Qualification and Validation).
  11. WHO Technical Report Series No. 986, Annex 2 – WHO Good Manufacturing Practices for Pharmaceutical Products: Main Principles.
  12. PIC/S PE 009 – Guide to Good Manufacturing Practice for Medicinal Products.
  13. PIC/S PI 041 – Good Practices for Data Management and Data Integrity in Regulated GMP/GDP Environments.
  14. ISPE GAMP 5 (Second Edition) – A Risk-Based Approach to Compliant GxP Computerized Systems.
  15. MHRA (2018) – GXP Data Integrity Guidance and Definitions.

Essential GMP Principles for Pharma Industry Professionals Read More »
Vendor Audit Questionnaire for Laminated Tube (Lamitube) Manufacturer

Vendor Audit Questionnaire for Laminated Tube: Below is a structured, regulator-aligned Vendor Audit Questionnaire tailored for a laminated tube (Lamitube) manufacturer supplying to pharmaceutical companies. The framework integrates expectations from World Health Organization, U.S. Food and Drug Administration, Pharmaceutical Inspection Co-operation Scheme, and European Medicines Agency guidelines (especially GMP for packaging materials, e.g., EU GMP Annex 1, WHO TRS 986/1019, FDA 21 CFR 210/211).


The questions are short, audit-friendly, and layered (Main → Sub-questions) so you can probe deeply during inspection.


🔷 1. Quality Management System (QMS)

Main Question:

How is the Quality Management System established, implemented, and maintained?

Breakdown:

  • Is there a documented Quality Manual aligned with GMP?
  • Are SOPs controlled, approved, and version-managed?
  • How is document lifecycle (issuance → archival) controlled?
  • Are quality objectives defined and periodically reviewed?
  • Is there a system for Quality Risk Management (ICH Q9 aligned)?
  • Are deviations, CAPA, and change control systems integrated?
  • Is management review conducted periodically?

🔷 2. Regulatory Compliance & Certifications

Main Question:

Does the company comply with applicable regulatory and GMP requirements?

Breakdown:

  • Which certifications are held (ISO 9001, ISO 15378, GMP)?
  • Are audits conducted by regulatory agencies or customers?
  • Are previous audit findings available with CAPA status?
  • Is there compliance with pharma packaging guidelines?
  • Are regulatory inspections documented and closed?

🔷 3. Personnel & Training

Main Question:

Are personnel qualified, trained, and compliant with GMP?

Breakdown:

  • Is there an organizational chart with defined responsibilities?
  • Are job descriptions documented?
  • How is initial and ongoing GMP training conducted?
  • Are training effectiveness evaluations performed?
  • Are personnel hygiene and gowning procedures defined?
  • Are operators trained in contamination control?

🔷 4. Premises & Facility Design

Main Question:

Is the facility designed to prevent contamination and mix-ups?

Breakdown:

  • Is material and personnel flow logically segregated?
  • Are clean/controlled areas defined (if applicable)?
  • Is there HVAC system qualification and monitoring?
  • Are pest control systems in place?
  • Are lighting, temperature, humidity controlled?
  • Are utilities (compressed air, water) monitored?

🔷 5. Equipment & Maintenance

Main Question:

Are equipment qualified, calibrated, and maintained?

Breakdown:

  • Is there a list of critical equipment?
  • Are IQ/OQ/PQ performed where applicable?
  • Is preventive maintenance planned and documented?
  • Are calibration records maintained?
  • Are breakdowns recorded and investigated?
  • Is cleaning of equipment validated or verified?

🔷 6. Raw Materials Control

Main Question:

How are raw materials (e.g., laminate, resins, inks) controlled?

Breakdown:

  • Are suppliers qualified and approved?
  • Are incoming materials tested against specifications?
  • Is there a quarantine → release system?
  • Are COAs verified?
  • Are storage conditions controlled?
  • Is FEFO/FIFO followed?

🔷 7. Production & Process Control

Main Question:

Are manufacturing processes controlled and validated?

Breakdown:

  • Are batch manufacturing records (BMR) available?
  • Are critical process parameters defined and monitored?
  • Is process validation performed?
  • Are line clearance procedures implemented?
  • Are in-process checks documented?
  • Is rejection/rework controlled?

🔷 8. Printing & Artwork Control (Critical for Lamitube)

Main Question:

How is printed artwork controlled to prevent errors?

Breakdown:

  • Is artwork approved by customers before printing?
  • Are master artworks securely controlled?
  • Are printing plates/cylinders controlled and verified?
  • Is line clearance done between different artworks?
  • Are barcode and text verification systems used?
  • Are reconciliation procedures in place?

🔷 9. Cleaning & Contamination Control

Main Question:

Are cleaning procedures effective to prevent contamination?

Breakdown:

  • Are cleaning SOPs defined for all areas?
  • Are cleaning records maintained?
  • Is cleaning validation required and performed?
  • Are residues (ink, adhesive) controlled?
  • Are cross-contamination risks assessed?

🔷 10. Laboratory Controls & Testing

Main Question:

Are QC laboratory operations reliable and compliant?

Breakdown:

  • Are specifications defined for tubes (dimension, leakage, print quality)?
  • Are test methods validated?
  • Is there an approved sampling plan?
  • Are instruments calibrated?
  • Are OOS (Out of Specification) handled properly?
  • Are retain samples stored?

🔷 11. Packaging & Finished Product Control

Main Question:

How are finished laminated tubes controlled and released?

Breakdown:

  • Are finished goods inspected before release?
  • Is batch traceability ensured?
  • Are packing materials controlled?
  • Are labeling and identification clear?
  • Is transport validation considered?

🔷 12. Storage & Distribution

Main Question:

Are storage and distribution conditions controlled?

Breakdown:

  • Are warehouses clean and organized?
  • Are environmental conditions monitored?
  • Are dispatch procedures documented?
  • Is traceability maintained during shipment?
  • Are damaged goods handled properly?

🔷 13. Deviations, CAPA & Change Control

Main Question:

Are quality issues effectively managed?

Breakdown:

  • Are deviations recorded and investigated?
  • Is root cause analysis performed?
  • Are CAPAs tracked for effectiveness?
  • Are changes evaluated for impact before implementation?
  • Is change control linked to validation?

🔷 14. Complaint Handling & Recall

Main Question:

Is there an effective system for complaints and recalls?

Breakdown:

  • Are customer complaints logged and investigated?
  • Is trending analysis performed?
  • Is there a mock recall system?
  • How quickly can products be traced and recalled?
  • Are regulatory authorities notified if required?

🔷 15. Data Integrity & Documentation

Main Question:

Is data integrity ensured across all systems?

Breakdown:

  • Are records attributable, legible, contemporaneous (ALCOA)?
  • Are electronic systems validated?
  • Are audit trails enabled?
  • Is access control implemented?
  • Are backups performed regularly?

🔷 16. Vendor & Supply Chain Control

Main Question:

How are upstream vendors controlled?

Breakdown:

  • Is there an approved vendor list (AVL)?
  • Are vendor audits conducted?
  • Are critical suppliers periodically re-evaluated?
  • Are agreements (Quality Agreements) in place?

🔷 17. EHS (Environment, Health & Safety)

Main Question:

Are safety and environmental controls implemented?

Breakdown:

  • Are MSDS available for chemicals (inks, solvents)?
  • Are hazardous materials stored properly?
  • Is waste disposal compliant?
  • Are fire safety systems installed?
  • Are emergency procedures defined?

🔷 18. Continuous Improvement

Main Question:

Does the company focus on continuous improvement?

Breakdown:

  • Are KPIs defined (rejection rate, complaints)?
  • Are internal audits conducted?
  • Are improvement projects documented?
  • Is trend analysis performed?

✅ Pro Tips for Strong Vendor Audit (Practical Execution)

  • Focus heavily on:
    • Artwork control (highest risk in tubes)
    • Traceability (batch → raw material → dispatch)
    • Line clearance & mix-up prevention
  • Ask for live records, not just SOPs.
  • Perform shop-floor verification (not just QA office review).
  • Cross-check:
    • BMR vs actual production
    • COA vs test raw data
  • Look for data integrity red flags (backdating, overwriting, missing logs)


🔴 SERIES–2: ADVANCED / FORENSIC VENDOR AUDIT QUESTIONS


🔷 1. QMS Effectiveness (Beyond Documentation)

Main Question:

How do you demonstrate that your QMS is effective, not just compliant?

Breakdown:

  • Show last 12 months trend of deviations vs CAPA closure time
  • What % of CAPAs are delayed? Justify
  • Provide example where CAPA failed → what was done next?
  • How do you ensure recurrence does NOT happen?
  • Are KPIs linked to management incentives?
  • Show evidence of risk-based decision making, not SOP-driven only

🔷 2. Data Integrity (Critical – High Risk Area)

Main Question:

How do you ensure ALCOA+ compliance in real operations?

Breakdown:

  • Show raw data vs reported COA → any mismatch?
  • Are there any blank spaces, overwriting, backdating in records?
  • Who reviews audit trails and how frequently?
  • Can operators delete or modify data?
  • Show one real audit trail for a batch
  • What controls prevent “testing into compliance”?
  • How do you detect unofficial records (shadow documentation)?

🔷 3. Deviation Investigation Depth

Main Question:

How robust is your root cause investigation?

Breakdown:

  • Show a major deviation report (last 6 months)
  • Was root cause scientifically justified or assumed?
  • Was 5 Why / Fishbone / FMEA used?
  • Were multiple root causes considered?
  • Was QA independent in conclusion approval?
  • How do you verify root cause correctness?

🔷 4. Change Control – Hidden Risks

Main Question:

How do you ensure changes do not introduce unseen risks?

Breakdown:

  • Show a recent change control affecting production
  • Was risk assessment formal (ICH Q9)?
  • Were validation/revalidation requirements evaluated?
  • Was customer notified (for critical changes)?
  • Were changes implemented before approval (backdoor change)?
  • How do you track temporary changes?

🔷 5. Process Validation – Real Assurance

Main Question:

How do you prove your process is consistently capable?

Breakdown:

  • Show Process Validation Report
  • Are worst-case conditions included?
  • What is the process capability (Cp/Cpk)?
  • How are critical parameters justified?
  • Is continued process verification (CPV) performed?
  • Show trend of in-process rejection over time

🔷 6. Artwork & Printing – Zero Error Expectation Area

Main Question:

How do you ensure zero artwork error, considering high regulatory risk?

Breakdown:

  • Show real case of artwork error → what happened?
  • Is there 100% vision inspection or manual?
  • How is mix-up between similar artworks prevented?
  • Are cylinders uniquely coded and verified?
  • Is there dual verification before printing?
  • What is reconciliation tolerance for printed tubes?

🔷 7. Line Clearance – Practical Effectiveness

Main Question:

How do you guarantee zero mix-up between batches/products?

Breakdown:

  • Demonstrate a live line clearance
  • Who verifies clearance (production vs QA)?
  • Are previous batch remnants physically checked?
  • Are labels/artwork fragments controlled?
  • Is photographic evidence used?

🔷 8. Supplier Risk Management

Main Question:

How do you control risks from your raw material suppliers?

Breakdown:

  • Show supplier risk categorization
  • How often are critical suppliers audited?
  • Any supplier disqualified recently? Why?
  • How do you verify supplier COA reliability?
  • Do you perform skip testing or full testing?

🔷 9. Laboratory Control – Reliability Check

Main Question:

How do you ensure QC results are scientifically reliable?

Breakdown:

  • Show OOS investigation (real case)
  • Was Phase I / Phase II investigation followed?
  • Are analysts trained on data integrity?
  • Any retesting without justification?
  • How are reference standards controlled?

🔷 10. Rejection, Rework & Scrap Control

Main Question:

How do you prevent rejected materials from re-entering the system?

Breakdown:

  • Show rejected batch records
  • Is rework scientifically justified and approved?
  • How is scrap physically destroyed?
  • Any case of unauthorized reprocessing?
  • Are rejection trends analyzed?

🔷 11. Traceability (Backward & Forward)

Main Question:

Can you trace any product within minutes?

Breakdown:

  • Perform mock traceability test:
    • Finished tube → raw material batch
    • Raw material → all affected customers
  • Time required to retrieve data?
  • Is system manual or ERP-based?
  • Any gaps in traceability observed?

🔷 12. Complaint Handling – True Root Cause

Main Question:

How do you ensure complaints lead to real improvement?

Breakdown:

  • Show last 5 complaints
  • Are trends analyzed (e.g., print defect, leakage)?
  • Were CAPAs effective?
  • Any repeated complaints for same issue?
  • Was customer feedback incorporated?

🔷 13. Recall System – Practical Readiness

Main Question:

How prepared are you for a real recall?

Breakdown:

  • Show last mock recall report
  • Time taken to complete recall simulation?
  • % traceability achieved?
  • Are customers informed in simulation?
  • Are regulatory requirements considered?

🔷 14. Cleaning Validation (Critical for Cross-Contamination)

Main Question:

How do you ensure no carryover of ink/chemical residues?

Breakdown:

  • Are worst-case products selected?
  • What is acceptance criteria (scientific or arbitrary)?
  • Are swab/recovery studies performed?
  • Is visual cleanliness justified scientifically?
  • Any failure observed in cleaning validation?

🔷 15. Environmental Control (Especially Printing Area)

Main Question:

How do environmental factors impact product quality?

Breakdown:

  • Are temp/humidity linked to print quality?
  • Any environmental excursions recorded?
  • Impact assessment done?
  • Are sensors calibrated?

🔷 16. Warehouse – Hidden GMP Gaps

Main Question:

How do you prevent mix-up and deterioration in storage?

Breakdown:

  • Are status labels (Quarantine/Released/Rejected) clear?
  • Any case of wrong material issuance?
  • Are returns segregated?
  • Is pest control effective (show logs)?

🔷 17. Internal Audit Effectiveness

Main Question:

Are internal audits truly identifying gaps?

Breakdown:

  • Show last internal audit report
  • Were critical findings raised?
  • Any repeat observations?
  • Is QA independent from production?

🔷 18. Management Commitment (Reality Check)

Main Question:

How does top management ensure GMP compliance?

Breakdown:

  • Are resources sufficient (manpower, budget)?
  • Any production vs quality conflict?
  • Example where batch was rejected despite pressure
  • Are quality metrics reviewed at top level?

🔥 HIGH-IMPACT AUDIT TECHNIQUES (Use During Audit)

  • Ask: “Show me now” instead of “Do you have”
  • Cross-question same topic with:
    • Operator
    • Supervisor
    • QA
  • Pick random batch → trace everything
  • Compare:
    • SOP vs actual practice
  • Watch for:
    • Delayed entries
    • Too-perfect records
    • Identical handwriting


🔴 SERIES–3: TRAP QUESTIONS (HIGH-IMPACT AUDIT TOOL)

⚠️ How to use:

  • Ask indirectly
  • Cross-check answers between departments
  • Always demand live evidence

🔷 1. “Show Me Now” Trap (Reality vs Documentation)

Main Trap Question:

“Can you show me a batch currently under processing and all associated live records?”

Trap Breakdown:

  • Does the operator hesitate or call QA first?
  • Are entries made in real-time or backfilled?
  • Do timestamps align with actual process time?
  • Is handwriting consistent or pre-filled?

👉 Red Flag: Records updated just before showing


🔷 2. Data Integrity Trap (Backdating Detection)

Main Trap Question:

“Pick any record from last week—can you explain when exactly this entry was made?”

Trap Breakdown:

  • Compare:
    • Ink color / pen variation
    • Time gaps between entries
  • Ask operator:
    • “Did you write this immediately or later?”

👉 Red Flag: Same pen, same handwriting for full shift data


🔷 3. Shadow Documentation Trap

Main Trap Question:

“Do operators ever use rough sheets before final entry?”

Trap Breakdown:

  • Check drawers, behind machines
  • Ask casually: “How do you remember readings before writing?”

👉 Red Flag: Unofficial notebooks / loose papers


🔷 4. Deviation Suppression Trap

Main Trap Question:

“In the last 3 months, did any process fail but was not recorded as deviation?”

Trap Breakdown:

  • Cross-check:
    • Maintenance logs
    • QC OOS
    • Production downtime
  • Compare with deviation register

👉 Red Flag: Events exist but no deviation raised


🔷 5. CAPA Effectiveness Trap

Main Trap Question:

“Show a deviation that happened twice—why did CAPA fail first time?”

Trap Breakdown:

  • Check recurrence
  • Evaluate depth of root cause

👉 Red Flag: Same issue repeating with different wording


🔷 6. Artwork Mix-up Trap (Critical for Tubes)

Main Trap Question:

“What prevents two similar artworks from being mixed during printing?”

Trap Breakdown:

  • Physically verify:
    • Cylinders
    • Printed tubes
  • Ask operator to explain difference

👉 Red Flag: Reliance only on visual check without system control


🔷 7. Line Clearance Trap (False Compliance)

Main Trap Question:

“Can we check the previous product run on this line?”

Trap Breakdown:

  • Look for:
    • Leftover labels
    • Tubes under machine
  • Ask: “Who verified clearance?”

👉 Red Flag: Clearance signed but physical evidence remains


🔷 8. Training Effectiveness Trap

Main Trap Question:

Ask operator directly:
“Explain what you will do if you find a defective tube?”

Trap Breakdown:

  • Compare with SOP
  • Check confidence level

👉 Red Flag: SOP exists but operator unaware


🔷 9. QC Testing Manipulation Trap

Main Trap Question:

“Have you ever repeated a test because the first result was not acceptable?”

Trap Breakdown:

  • Ask follow-up:
    • “Was it documented?”
  • Check raw data vs final COA

👉 Red Flag: Retesting without investigation


🔷 10. OOS Handling Trap

Main Trap Question:

“Show me an OOS result and explain full investigation”

Trap Breakdown:

  • Check:
    • Hypothesis-driven investigation
    • Lab error justification

👉 Red Flag: Blaming analyst without proof


🔷 11. Change Control Bypass Trap

Main Trap Question:

“Have you ever implemented a change urgently before approval?”

Trap Breakdown:

  • Cross-check with production/engineering
  • Verify temporary changes

👉 Red Flag: Verbal approvals / undocumented changes


🔷 12. Supplier COA Trust Trap

Main Trap Question:

“Do you always test raw materials or rely on supplier COA?”

Trap Breakdown:

  • Ask:
    • “When was last full testing done?”

👉 Red Flag: Blind trust in supplier without verification


🔷 13. Traceability Stress Test (Live Challenge)

Main Trap Question:

“Trace this finished tube to raw materials within 10 minutes”

Trap Breakdown:

  • Observe:
    • Time taken
    • Data gaps

👉 Red Flag: Delays, incomplete traceability


🔷 14. Scrap Control Trap

Main Trap Question:

“What happens to rejected printed tubes?”

Trap Breakdown:

  • Physically inspect scrap area
  • Check destruction records

👉 Red Flag: Scrap accessible / reusable


🔷 15. Warehouse Mix-up Trap

Main Trap Question:

“Can you show any wrongly placed material found in last audit?”

Trap Breakdown:

  • Check labeling system
  • Verify segregation

👉 Red Flag: Same-looking materials stored together


🔷 16. Environmental Excursion Trap

Main Trap Question:

“What happens if humidity exceeds limit during printing?”

Trap Breakdown:

  • Ask for real case
  • Check deviation link

👉 Red Flag: No impact assessment


🔷 17. Internal Audit Integrity Trap

Main Trap Question:

“Why were no critical observations found in last internal audit?”

Trap Breakdown:

  • Compare with your findings

👉 Red Flag: Internal audit too “clean”


🔷 18. Management Pressure Trap

Main Trap Question:

“Has production ever continued despite quality concern?”

Trap Breakdown:

  • Ask different levels separately

👉 Red Flag: Conflicting answers


🔷 19. “Too Perfect System” Trap

Main Trap Question:

“You have zero deviations last month—how is that possible?”

👉 Red Flag: Indicates underreporting


🔷 20. Contradiction Trap (Powerful Technique)

Method:

Ask same question to:

  • Operator
  • QA
  • Supervisor

👉 Compare answers

Example:

  • “Who performs line clearance?”
  • “Who approves deviation?”

👉 Red Flag: Different answers


🔥 CRITICAL OBSERVATION SKILLS (MOST IMPORTANT)

During audit, continuously watch for:

  • ⛔ Entries written in same handwriting
  • ⛔ No corrections in months (unrealistic)
  • ⛔ Fearful operators
  • ⛔ QA answering everything (production silent)
  • ⛔ Delayed data retrieval
  • ⛔ “We always follow SOP” without explanation

🧠 PRO AUDITOR STRATEGY (FIELD-LEVEL)

  • Never rely on verbal answers
  • Always:
    • See → Verify → Cross-check
  • Use silence pressure (wait after asking)
  • Randomly select:
    • Batch
    • Operator
    • Record

✅ If you want FINAL LEVEL (ELITE)

I can prepare:

🔷 “Audit Scoring System”

  • Critical / Major / Minor classification
  • Risk-based scoring (FDA-style)

🔷 “Lamitube-Specific Risk Matrix”

  • Leakage risk
  • Print toxicity risk
  • Extractables/leachables concern

🔷 “DGDA Bangladesh Audit Angle”

  • Local regulatory expectations
  • Common inspection findings in Bangladesh

Vendor Audit Questionnaire for Laminated Tube (Lamitube) Manufacturer Read More »

What is a Fishbone Diagram, how is it used?

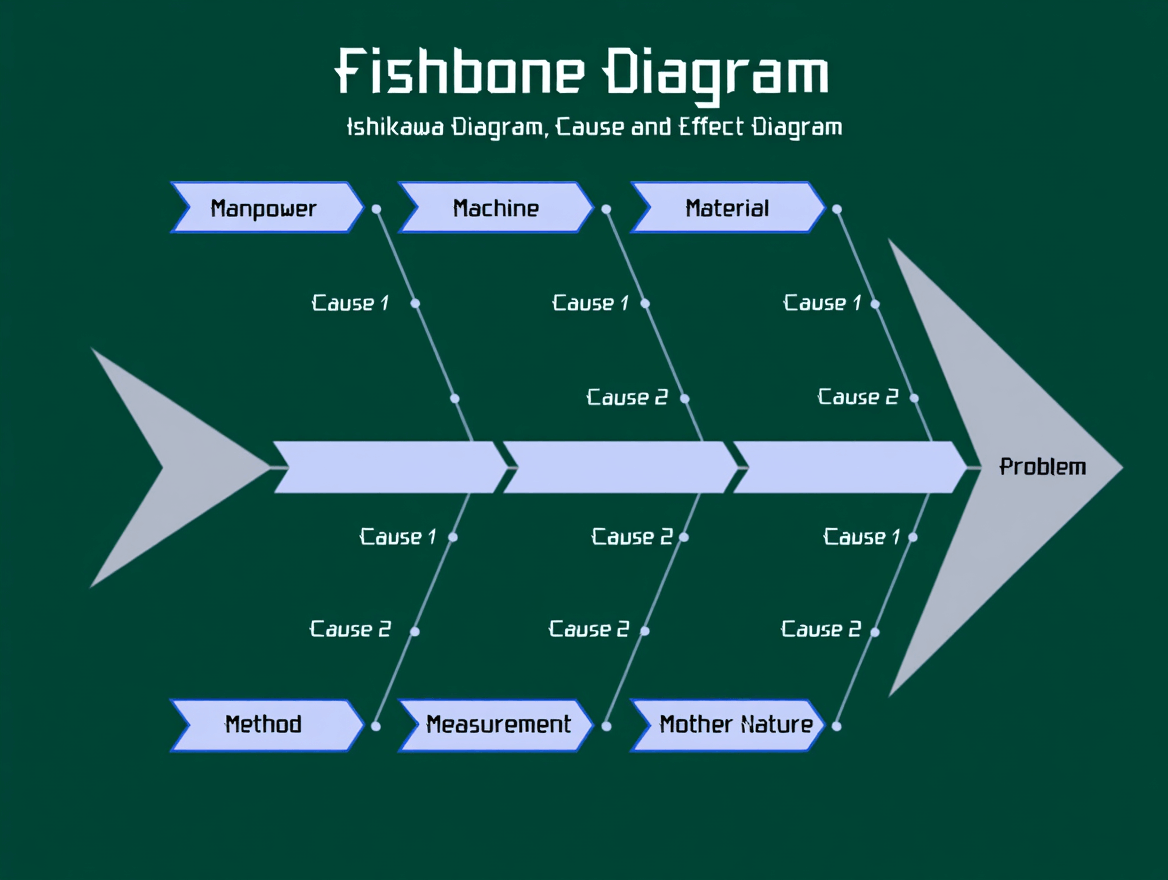
A Fishbone Diagram (also called Ishikawa Diagram or Cause-and-Effect Diagram) is a visual tool used to systematically identify and organize all possible causes of a specific problem or effect. It was developed by Dr. Kaoru Ishikawa in the 1960s.

The diagram looks like a fish skeleton:

  • The “head” of the fish is the problem/effect you want to solve.
  • The “bones” branching off the spine are the major categories of causes.
  • Smaller bones branching off are the specific causes within each category.

Why use it?

  • Forces teams to think through all possible causes (not just the obvious ones)
  • Prevents jumping to conclusions
  • Encourages cross-functional collaboration
  • Very useful in root cause analysis (RCA), quality improvement, Six Sigma, Lean, etc.

How to Create and Use a Fishbone Diagram (Step-by-Step)

  1. Define the problem clearly
    Write it in a box on the right (the “head”). Be specific.
    Example: “Customer complaints about late deliveries increased by 40% in Q3”
  2. Draw the main spine
    A horizontal arrow pointing to the problem.
  3. Identify major cause categories (the big bones)
    Common categories (the 6 Ms for manufacturing, or adapt to your industry):

    • Man (People)
    • Method (Processes)
    • Machine (Equipment/Technology)
    • Material
    • Measurement
    • Mother Nature (Environment)

    For service/industry, people often use:

    • People, Policies, Procedures, Plant/Technology, etc.
  4. Brainstorm all possible causes
    Ask “Why does this happen?” repeatedly (5 Whys technique helps).
    Write each cause as a branch off the relevant category.
  5. Go deeper
    For each cause, ask “Why?” again and add sub-causes (smaller bones).
  6. Analyze and prioritize
    Circle the most likely root causes (use voting, data, Pareto, etc.).
  7. Develop action plan
    Address the confirmed root causes.

Practical Case Study: Restaurant Getting Complaints About Cold Food

Problem (Head of the fish):
“Customers frequently complain that food arrives cold” (complaints rose from 3 to 18 per week)

Fishbone Diagram Categories Used:
We used 6 categories suitable for a restaurant:

  1. People (Staff)
    • Servers forget to check food temperature before leaving kitchen
    • New staff not trained on urgency
    • Kitchen and serving staff not communicating
    • Too few servers during peak hours → delay
  2. Processes (Methods)
    • No standard procedure for checking food temperature
    • Food waits too long on the pass before being served
    • Tickets not prioritized correctly during rush
    • No “food ready” notification system between kitchen and floor
  3. Equipment (Machines)
    • Heat lamps broken or insufficient
    • Plates not pre-heated
    • Delivery trays too small → food stacked and cools faster
    • Old warming drawers not maintaining temperature
  4. Materials
    • Some dishes (e.g., pasta) cool faster than others
    • Large portion sizes take longer to eat → perceived as cold
  5. Environment (Mother Nature/Place)
    • Dining room AC set too low near some tables
    • Long walking distance from kitchen to farthest tables
    • Draft from entrance door
  6. Measurement (or Management)
    • No tracking of time from “food ready” to “delivered to table”
    • No temperature checks logged
    • No customer feedback analyzed by dish/table location

Root Causes Identified After Investigation (circled on the diagram):

  • Heat lamps were broken for 3 weeks (maintenance backlog)
  • No policy to pre-heat plates
  • Average time from kitchen to table was 4.5 minutes during peak (target should be <2 min)
  • Servers overloaded: 1 server handling 9 tables instead of max 6

Actions Taken:

  1. Repaired/replaced heat lamps same week
  2. Implemented plate pre-heating in salamander broiler
  3. Added a runner position during peak hours
  4. Created a simple checklist: food temperature >60°C before leaving pass
  5. Installed digital ticket system with timers

Result:
Cold food complaints dropped from 18 to 2 per week within one month.

What is a Fishbone Diagram, how is it used? Read More »

Risk Assessment Procedure

Risk Assessment , Purpose :

Risk Assessment , The objective of carrying out the Risk Assessment is to ensure the potential threats to the product quality/ supply are properly assessed and appropriate control measures are in place.

Risk Assessment , Scope :

This procedure applies for the risk assessment of facilities, utility services, equipment & machinery, warehousing, manufacturing, packaging, testing and transfer of quality products from at XX Pharmaceuticals Limited (Both General and Sterile Block).

Definitions / Abbreviation:

[][]Risk assessment: Risk assessment consists of the identification of product quality risk and evaluation of risks. Quality risk assessments begin with a well-defined problem description or risk question. When the risk in question is well defined, an appropriate risk management tool and the types of information needed to address the risk question will be more readily identifiable.

Responsibilities:

[][]The roles and responsibility is as follows

All concerned department heads

[][]To perform the risk assessment associated in his area in a cross-functional team, including one QA personnel as a mandatory member.

Manager, Quality Assurance

[][]To ensure that proper risk assessment is done if there any potential risk(s) involved in the product quality/ supply.

Head of Quality Assurance

[][]Approval of the SOP.
[][]Implementation of risk assessment procedure

Procedure:

=>Risk assessment: Risk assessment consists of the identification of product quality risk and evaluation of risks. Quality risk assessments begin with a well-defined problem description or risk question. When the risk in question is well defined, an appropriate risk management tool and the types of information needed to address the risk question will be more readily identifiable. As an aid to clearly defining the risk(s) for risk assessment purposes, three fundamental questions are often helpful:
[][]What might go wrong?
=>Output for Question 1: Problem/ Failure descriptions- Cause – Failure mode – Effects
[][]What is the likelihood (probability) it will go wrong?
[][]What are the consequences (severity)?
=>Output for Question 2 & 3: Risk Class, Job/Action Priority, Failure rates, Risk priority number (RPN).

[][]Risk identification: Risk identification is a systematic use of information to identify product quality risk or problem description. Information can include historical data, theoretical analysis, informed opinions, and the concerns of stakeholders.

[][]Risk analysis: Risk analysis is the estimation of the risk associated with the product quality. It is the qualitative or quantitative process of linking the likelihood of occurrence and severity of harms. In some risk management tools, the ability to detect the harm (detectability) also factors in the estimation of risk.

[][]Risk evaluation: Risk evaluation compares the identified and analyzed risk against given risk criteria. Risk evaluations consider the strength of evidence for all three of the fundamental questions.

[][]The output of a risk assessment is either a quantitative estimate of risk or a qualitative description of a range of risk. When risk is expressed quantitatively, a numerical probability is used.

[][]Alternatively, risk can be expressed using qualitative descriptors, such as “high”, “medium”, or “low” (Risk class one, Risk class two or Risk class three) , which should be defined in as much detail as possible.

[][]Sometimes a “risk score” is used to further define descriptors in risk ranking. In quantitative risk assessments, a risk estimate provides the likelihood of a specific consequence, given a set of risk-generating circumstances.

[][]Thus, quantitative risk estimation is useful for one particular consequence at a time.

[][]To access and manage risk the following recognized tools will be applied:
=>Risk ranking and filtering will be applied for qualitative (High, Medium or low) risk analysis.
=>Failure Mode Effect Analysis (FMEA) will be used for quantitative estimate of risk. (Score of RPN).
=>Fish Bone Diagram/ Cause & Effect Diagram Analysis

Procedure for Risk ranking and filtering:

[][]Identify the key stages for the part of the supply chain for which the company has responsibility from material supply through to use of product by patients & customers.
[][]Identify potential threats that could impact the product quality or security at each key stage.
[][]Estimate the consequences both in terms of threats and opportunities may be high, medium or low.

[][]Evaluate the exiting control measures for their effectiveness at controlling the threats.
[][]Determine the risk to product quality associated with each threat.
[][]Establish a plan for the introduction of improved controls (additional control measures) where existing threats are not adequately addressed.
[][]Implement the additional control measures and monitor their effectiveness.
[][]To identify the threats the following points should be considered:
=>Patient : Hazard/ risk to the patient safety, patient compliance.
=>Personnel : Attributes, training, education, competence, communication
=>Equipment : Type, design, condition, capacity, location, installation, operation, maintenance & calibration.
=>Facility : Layout, utilities, maintenance, dedication & hygiene.
=>Methods & Procedures : Checking, content, alterations, distribution, utilization, condition, change control, storage, trends, handling planned & abnormal events.
=>Materials : Identity, status, control, quantity, handling, specification, security arrangements, counterfeiting controls & material condition.
=>Environment : Physical effects of climatic & storage conditions (temperature, time, humidity, rain, air pressure, light, vibration etc.), pest infestation, contamination, and damage due to fire or natural disaster like flood, tornado, earthquake etc.

[][]When threats have been identified and commented upon, the table in Annexure – I should be completed.
[][]A judgment should be made on the severity of the consequences & the probability/ likelihood of the adverse events occurring taking into consideration any current control measures that are in place. Each of this can be “low”, “medium”, or “high”.
[][]Considering the severity and probability/likelihood of the events risk will be expressed Risk class one, Risk class two or Risk class three (or, High risk”, “medium risk”, or “low risk).
[][]Risk filtering will be done considering the Risk class (i.e. Risk class one, Risk class two or Risk class three) and detection of the events. Jobs/ actions will be prioritized as High priority, Medium priority or Low priority.
[][]All the jobs/actions for the changed situation shall be identified with in-depth analysis making sure all the impacted areas and the actions with the documentation requirements are assessed and completion date against all the identified actions are set.
[][]Below are two matrixes for clear understanding.

Procedure for Failure Mode Effect Analysis (FMEA):

[][]Risk in an FMEA evaluation has three components: Severity, Probability and detection/ detectability. The first step in any risk assessment is to define the component of FMEA
[][]Definition of the components of the FMEA are:
[][]Severity: if a failure were to occur, what effect would that failure have on the product quality and on the patient (if any)?
[][]Probability of occurrence: how likely is it for a particular failure to occur?
[][]Detectability (ability to detect): what mechanisms are in place (if any) to detect a failure if it were occur?
[][]Each of the above components requires clear descriptions and a corresponding scale to rank or score the projected impact (i.e. a scale for Severity; a scale for Probability; and a scale for ability to Detect). In addition, a composite score would then need to be calculated (e.g. severity multiplied by Probability multiplied by ability to detect)
[][]Non sequential number (e.g. 1,3,5,7, 9) will be used for probability and detection as the use of non-consecutive numbers allow more distinction between rating (table 2 & table 3) and to put more emphasis on the severity criteria a non-linear scoring scale will be utilized (e.g. 1, 4, 9,16, 25) . Please see table 1 for details.

[][]Table 1: Severity criteria for FMEA

Severity
ValueDescriptionCriteria
1IrrelevantNo impact to product quality and process robustness
4SlightNo impact to product quality
9ImportantNoticeable impact to product quality, but can be recovered by reprocessing
16CriticalDefinite impact to product quality that may require rework
25DisastrousBatch failure, not recoverable by rework

Note: Criteria in the above table will be changed based on the subject under assessment

[][]Table 2: Probability criteria for FMEA

Probability
ValueDescriptionCriteria
1An unlikely probability of occurrenceFailure has never been seen in any relevant lab experiments, or scale-up batches yet but it is theoretically possible.
3A remote probability of occurrenceFailure only seen once or twice in relevant lab experiments, never in scale-up batches.
5An occasional probability of occurrenceFailure potential has been noted in several relevant lab experiments, or at scale-up. If procedures are followed the failure potential is minimal.
7A moderate probability of occurrenceFailure potential has been noted in several relevant lab experiments, or at scale-up, in-process control maybe required to avoid failure.
9A high probability of occurrenceFailure potential has been noted in several relevant lab experiment, or at scale-up, an active non-standard feedback control loop may be required.

Note: Criteria in the above table will be changed based on the subject under assessment

[][]Table 3: Detectability criteria for FMEA

Detection
ValueDescriptionCriteria
1High degree of detectabilityA: Validated automatic detection system that is a direct measure of failure.
B: Two or more manual operated validated detection systems, direct or indirect. (e.g. Control range and IPC)
3Good detectabilityA: Single manually operated validated detection system that is a direct measure of failure. (e.g. IPC of failure, validated PAT)
5Likely to detectA: Single manually operated validated detection system that is not a direct measure of failure.
(e.g. PAT measurements or IPC's not directly linked to failure)
7Fair detectabilityA: Non validated (manual or automated) detection.
(e.g. visual level check, visual inspection of vessels).
9Low or no detectabilityNo ability to detect the failure

Note: Criteria in the above table will be changed based on the subject under assessment

Risk Scoring Matrix

[][]The composite risk score for each unit operation step is the product of its three individual component ratings: severity, probability, and detection. This composite risk is called a risk priority number (RPN).
RPN = S x P x D (S= Severity; P= Probability & D= Detection)
[][]The RPN number is not absolute and should be considered in context with other factors that influence the product risk outside the scope of this evaluation. The RPN provides a relative priority for taking action – the bigger the RPN, the more important to address the corresponding failure being assessed.

[][]The table in Annexure – III will be used for FMEA. For each formulation component or manufacturing processing step under evaluation, the function of the component or processing step, potential failure mode and effect of the failure mode should be recorded.

[][]A severity score is then assigned. The root cause of the failure is described and a score is assigned to the probability of occurrence of the failure. Controls that are currently in place to detect the failure are listed and a detection score is then assigned.

[][]The RPN number is calculated. The action(s) that need to be taken to reduce or mitigate the risk are listed and individuals or departments responsible for implementing the actions are identified with target dates for completion.

[][]All the actions for the changed situation shall be identified with in-depth analysis making sure all the impacted areas and the actions with the documentation requirements are assessed and completion date against all the identified actions are set.

Fish Bone Diagram/ Cause & Effect Diagram Analysis

[][]The root cause analysis tool used for major or critical deviation, OOS, market complaint to identify quality defect prevention and potential factors causing an overall effect. Each cause or reason for imperfection is a source of variation. Causes are usually grouped into major categories to identify these sources of variation.

Defining “Effect”

[][]The first step in using the fishbone diagram as a problem solving tool is to clearly define your effect, or outcome that you don’t like. This could be a quality issues, not meeting metrics or troubleshooting the introduction of a new process or product line. This becomes the “head” of the diagram. Use butchers paper or a whiteboard to sketch out the fishbones template.

[][]Defining an effect takes a little practice. Make sure it is brief and succinct. Use facts and numbers where possible. Spend a few minutes reflecting on your effect with the team; does everyone agree that the statement defines the problem as fully as possible?
Brainstorming the “Causes”
[][]With your team, we want to add the bones to this diagram, brainstorming all of the possible influencing factors. Each idea needs to be put into a category or branch.
[][]The following probable categories to be assessed the probable causes through brainstorming is also known as 6M as per annexure-IV
=>Man
=>Machine
=>Method
=>Measurement
=>Material
=>Mother Nature

[][]Man/People/Personnel: Everyone involved with the process across the value stream, including support functions Processes / Methods: This defines how the process is performed and the all requirements needed for doing it, including quality procedures, work orders / travelers / work instructions, drawings.
[][]Machines / Equipment: All machines and equipment, needed to accomplish the job, including tools.
[][]Materials: Raw & Packaging materials, purchased parts and sub-assemblies that feed into the end product.
[][]Measurements: defines how have we determined that the outcome is wrong.

[][]Mother Nature: The standard one which turns to wrong or deviation of the standard outcome
[][]As the team suggests possible causes, determine which heading that idea belongs under, jotting it down clearly. Also add another branch, covering “why” that cause would influence the effect we are investigating. Continue until the team runs out of ideas.

[][]If is there any branches of the diagram that are missing, develop into that area further, asking questions; “Is it possible that the environment has affected our problem” too hot, too cold, too wet?
Document Numbering
[][]The unique document numbering for Risk Assessment shall consist of 9 (nine) alpha-neumerical characters, broken down as follows –
=>e.g. RA/XXX/YY
Where,
=>RA is the Risk Assessment
=>/ is separator
[][]XXX is the sequential number starting from 001, 002 & so on for a calendar year which will be further start from 001 for the next year.
=>YY is the last two digits of the year
=>For example; RA/001/YY
=>Here RA means Risk Assessment
=>001 is the sequential number
=>YY represents for the year 20YY
[][]QA shall issue the Risk Assessment document number and maintain the log register (Annexure – II).

Annexure:

Annexure-I: Risk Assessment Table and Action Plan
Annexure-II: Log Register for Risk Assessment
Annexure-III: Risk Assessment Table and Action Plan
Annexure-IV: Fish Bone Diagram Chart

Risk Assessment Procedure Read More »

Floor Inspection by QA Inspector

Floor Inspection, Purpose :

Floor Inspection, The purpose of this SOP is to describe the roles, responsibilities and activities of production and QC floor inspection and IPCs check by QA personnel and to ensure Assurance with cGMP requirements.

Floor Inspection, Scope :

This procedure is applicable for cGMP observations of production and QC floor at XX Pharmaceuticals Limited (Both General and Sterile Block).

Definitions / Abbreviation:

[][]IPC : In Process Check
[][]QA : Quality Assurance
[][]QC : Quality Control
[][]OOS : Out Of Specification
[][]DT : Disintegration Time

Responsibilities:

[][]The roles and responsibility is as follows:

Quality Assurance Personnel

[][]Responsible for inspection of Production and QC floor to find out any cGMP observations.

Manager, Quality Assurance

[][]To ensure implementation of the procedure

Head of Quality Assurance

[][]Approval of SOP

Procedure:

[][]To assure Assurance with specification throughout a production process, Production Department will conduct the IPC checks following the In-process checks procedure.
[][]IPC inspectors of QA will verify the IPC data independently.
[][]The IPC specification will be mentioned in the relevant product specification and BMR & BPR.
[][]All OOS results will be investigated as per OOS results procedure.
[][]IPC inspector will verify the IPC data based on the frequency described in Annexure–I.
[][]During manufacturing of a batch, production personnel will check the IPC parameters.
[][]If a IPC data goes outside specification, production IPC checking personnel and QA IPC inspector will inform to production Executive for corrective measures.
[][]If QA IPC inspector observes that there is frequent failure of the IPC results then QA IPC inspector will notify the issue to Departmental Manager and Manager, Quality Assurance.

All IPCs will be performed by trained and certified QCOM and Production personnel. Inspection of

=>Adherence to Line clearance
=>Environmental conditions
=>Log books
=>General house keeping
=>Labelling status
=>Sampling
=>Reconciliation
=>Calibration status
=>Deviation
=>Pest controls
=>Cleaning & Contamination
=>Change Control
=>Document management
=>Adherence to maintenance schedule

Inspect GMP compliance in

=>Storage of Raw/Packaging materials/Finished packs
=>Manufacturing/packaging/cleaning operations
=>Dispensing of materials
=>Compliance with SOPs
=>Daily balance monitoring records
QA inspectors will visit the production floor daily. The inspection applies to all operations, specially The following:
=>If any unusual observation comes to the notice of area QA inspector, he will immediately communicate to the Departmental Executive. This report will be recorded in Daily IPC report format (Annexure-II).
=>This inspection report will be forwarded to Manager, Quality Assurance for further action.
=>QA inspectors will collect retention samples of finished packs from the running Secondary packaging line belts & record sampling date and quantity sampled in BPR with sign & date as per Sampling SOP.

In-Process Controls Check:

[][]QA Inspectors will independently conduct all In-process checks in addition to that done by production personnel as Procedure for In-Process Checks.
[][]QA inspectors will conduct the following IPC checks as per frequency described in Annexure–I and as a minimum at least once from every batch of product. This should be done at the start of every batch:

[][]Product: Tablets/Capsule

=>Manufacturing Stage: Encapsulation/ Compression/Coating
=>IPC Checks: Appearance, Average weight, Uniformity of weight, Hardness, Thickness, DT, RH & Temperature
=>Manufacturing Stage: Blister Packing
=>IPC Checks: Leak test, Overprinting, Cutting, RH, Temperature, Product name, Strength, Product code, Mfg & Expiry date, Price and Status label etc.

[][]Product: Powder for Suspension

=>Manufacturing Stage: Manufacturing, Filling & Packing
=>IPC Checks: Appearance, Uniformity of weight, Leak Test, RH, Temperature, Product code, Mfg & Expiry date, Overprinting, Price and Status label etc.

[][]Product: Sterile (Capsules/ Tablets)

=>Manufacturing Stage: Encapsulation/ Compression/Coating
=>IPC Checks: Appearance, Average weight, Uniformity of weight, Hardness, Thickness, DT, RH & Temperature.
=>Manufacturing Stage: Blister Packing
=>IPC Checks: Leak Test, Overprinting, Cutting, RH, Temperature, Product code, Mfg & Expiry date, Price and Status label etc.

[][]Product: Sterile (Powder for Suspension )

=>Manufacturing Stage: Manufacturing, Filling & Packing
=>IPC Checks: Appearance, Uniformity of weight, Leak Test, RH, Temperature, Product code, Mfg. & Expiry date, Overprinting, Price and Status Label etc.

[][]Product: Dialysis Fluid

=>Manufacturing Stage: Manufacturing, Filling & Packing
=>IPC Checks: A Appearance, pH, Weight, Induction Sealing, Cap Sealing, Leak Test, Temperature, Pressure

[][]QA inspectors will verify the IPC results as per specification.
[][]If any batch is completed before inspection of QA IPC inspector, then again QA IPC inspectors will verify the some critical parameters.
[][]In-process test failures must be brought to the attention of the Departmental Executive/ Manager & also Manager, Quality Assurance and appropriate action shall be taken and recorded.
[][]Any problem identified at production floor during printing or in-process checking during packaging, Problem observer immediately inform it, to his/her supervisor and if required Online Problem Notification Form (Annexure-III) to be raised by production through QA.
[][]Reference No. for Online Problem Notification will be as
PN-001/02/XX
Where-
=>PN represents Problem Notification
=>001 represents sequential number
=>02 represent Month of February
=>XX represents year of 20XX

[][]If any receiving quantity of packaging material requires replacement to run production smoothly, it shall be raised by production with Material Replacement Form (Annexure-IV) to warehouse through QA with justified reason for replacement.
[][]Reference No. for Material Replacement will be as
MR-001/02/XX
Where-
=>MR represents Material Replacement
=>001 represents sequential number
=>02 represent Month of February
=>XX represents year of 20XX
[][]The copy of raised Online Problem Notification Form should be sent to Manager, Supply Chain & Management with recommendation of Head of Quality Assurance if required.

QC Floor Inspection:

[][]QA IPC Inspector will also visit the Quality Control laboratory once daily for GLP in place and in use check.

Annexure:

Annexure-I: In-Process Checks
Annexure-II: Daily IPC Finding and Observation Report Sheet
Annexure-III: Online Problem Notification Form
Annexure-IV: Material Replacement Form

Floor Inspection by QA Inspector Read More »