Quality Assurance

Building a Strong Quality Management System

Dr. Sarah Ahmed

Karachi, Pakistan


Introduction

Every organization promises something to its customers: a product that works, a service that arrives on time, a result that can be trusted. A Quality Management System (QMS) is how an organization turns that promise into a repeatable reality. It is the set of policies, processes, responsibilities, and records that direct and control how an organization meets customer and regulatory requirements and improves over time.

Many organizations treat quality as a department, a binder of procedures, or a certificate on the wall. Those organizations often find that quality problems keep returning no matter how many procedures they write. A strong QMS is different. It is woven into daily work, supported by leadership, and built around learning from data and mistakes.

This article explains what makes a QMS strong, the core building blocks, a practical path to implementation, and the pitfalls that commonly undermine even well-intentioned efforts.


1. What a Quality Management System Really Is

A QMS is not a single document or software tool. It is an integrated system that answers a few fundamental questions:

  • What do our customers and regulators require of us?
  • What processes must we run to meet those requirements?
  • Who is responsible for each process, and how do we know it is working?
  • What do we do when something goes wrong?
  • How do we get better over time?

The international standard ISO 9001:2015 describes a QMS framework that can be applied to any organization, regardless of size or sector. Regulated industries add their own requirements on top. Medical device manufacturers follow ISO 13485, pharmaceutical manufacturers follow Good Manufacturing Practice and ICH Q10, and other sectors such as aerospace and automotive have their own standards. The underlying logic across all of them is the same: define your processes, control them, measure them, and improve them.


2. The Seven Quality Management Principles

ISO 9000:2015 and ISO 9001:2015 rest on seven principles that serve as the philosophical foundation of a strong QMS:

  1. Customer focus. The primary aim is to meet customer requirements and strive to exceed expectations. Every other principle serves this one.
  2. Leadership. Leaders establish unity of purpose and direction and create conditions in which people are engaged in achieving quality objectives.
  3. Engagement of people. Competent, empowered, and engaged people at all levels are essential to creating and delivering value.
  4. Process approach. Consistent and predictable results are achieved when activities are understood and managed as interrelated processes that function as a coherent system.
  5. Improvement. Successful organizations have an ongoing focus on improvement.
  6. Evidence-based decision making. Decisions based on the analysis and evaluation of data are more likely to produce desired results.
  7. Relationship management. Organizations manage their relationships with interested parties, such as suppliers, to sustain success.

These are not slogans. Each one translates into specific system requirements, and a QMS that ignores any of them tends to develop weak spots. For example, a system with excellent procedures but disengaged leadership will stall, and a system with strong leadership but poor data will make decisions on guesswork.


3. Leadership: The Foundation of Everything

The single most important factor in a strong QMS is top management commitment. Quality systems fail most often not because the procedures are flawed but because leadership treats quality as someone else’s job.

Effective leaders do several things:

  • Set a clear quality policy that reflects the organization’s purpose and is communicated and understood at every level.
  • Establish measurable quality objectives aligned with business strategy, such as reducing customer complaints by a defined percentage or improving on-time delivery.
  • Provide resources, including people, time, training, equipment, and technology.
  • Assign clear roles and authorities, including a quality function with enough independence and standing to raise concerns without fear.
  • Participate in management review, using the meeting to make real decisions rather than rubber-stamp reports.
  • Model the behavior they expect. If leaders bypass procedures under schedule pressure, employees will learn that quality is optional.

A useful test is to ask what happens when quality conflicts with a shipment deadline. The answer reveals the true priorities of the organization, regardless of what the policy says.


4. Understanding Context, Stakeholders, and Scope

A QMS should be designed for the organization it serves rather than copied from a template. ISO 9001 asks organizations to begin by understanding their context: the internal and external factors that affect their ability to deliver quality results. These include market conditions, technology, regulatory changes, organizational culture, and resource constraints.

Organizations must also identify interested parties (customers, regulators, employees, suppliers, shareholders, and the community) and determine which of their needs and expectations become requirements. This analysis then defines the scope of the QMS, meaning which products, services, sites, and processes it covers.

Skipping this step is a common mistake. A scope that is too narrow leaves important risks unmanaged, and a scope that is too broad creates bureaucracy without benefit. Time spent here saves rework later.


5. The Process Approach

A strong QMS treats the organization as a network of processes, each with inputs, activities, outputs, owners, and performance measures. Rather than organizing quality around departments, the process approach follows work as it flows from customer need to customer satisfaction.

Building a process-based system typically involves:

  • Mapping core processes, such as design and development, purchasing, production or service delivery, inspection and testing, and complaint handling.
  • Identifying support processes, such as training, maintenance, calibration, document control, and information technology.
  • Defining interactions, meaning where the output of one process becomes the input of another.
  • Assigning process owners who are accountable for performance and improvement.
  • Setting process metrics, such as yield, defect rate, cycle time, or first-pass success.

Process maps, flowcharts, and SIPOC diagrams (Suppliers, Inputs, Process, Outputs, Customers) are helpful tools. The goal is clarity: anyone should be able to see how work moves through the organization and where quality is won or lost.


6. Risk-Based Thinking

Modern quality standards put risk at the center of the system. ISO 9001:2015 builds risk-based thinking into planning, and ICH Q9 offers a detailed framework for Quality Risk Management in pharmaceutical settings. The idea is simple: anticipate what could go wrong and focus effort where the consequences would be greatest.

A practical risk approach includes:

  • Identifying risks and opportunities at the organizational, process, and product levels.
  • Assessing the likelihood and severity of each, often using a scoring matrix.
  • Prioritizing and deciding whether to eliminate, reduce, transfer, or accept each risk.
  • Planning actions proportionate to the risk, then integrating them into the QMS.
  • Reviewing risks regularly, because they change as products, suppliers, and markets change.

Tools such as FMEA (Failure Mode and Effects Analysis) and HACCP help structure this thinking. A well-run risk process prevents the trap of treating every issue as equally urgent, which wastes resources and dilutes attention from what matters most.


7. Documented Information and Document Control

Documentation is often what people picture when they think of a QMS, and for good reason. Documents communicate requirements, ensure consistency, and provide evidence that work was done as intended. But more documentation does not mean a stronger system.

A strong approach follows these principles:

  • Document what adds value. Write procedures where consistency matters, where errors are costly, or where regulations require them.
  • Keep them usable. Procedures should be short, clear, and written in language that operators actually use. Visuals, checklists, and work instructions often work better than long prose.
  • Control versions. Only current, approved documents should be available at points of use, and obsolete ones should be withdrawn.
  • Maintain records. Records such as inspection results, training files, calibration certificates, and audit reports demonstrate that the system is working.
  • Protect integrity. Records should be accurate, legible, secure, and retrievable for their required retention period. In regulated industries, electronic records need validated systems, access control, and audit trails.

A good test of documentation quality is whether the people doing the work helped write it and can follow it without a supervisor explaining it.


8. People, Competence, and Culture

Even the best-designed system depends on the people who run it. Building competence and culture is therefore central.

Competence. Organizations should define the skills and knowledge each role requires, assess gaps, provide training, and evaluate whether the training worked. Attendance at a session is not proof of competence. Observation, testing, and performance data are better indicators.

Awareness. Employees should understand the quality policy, how their work affects quality objectives, and the consequences of not following requirements.

Communication. Information must flow in all directions. Frontline staff often see problems first, so there must be safe and simple ways to raise them.

Culture. A strong quality culture is one where people take ownership, report errors openly, and focus on fixing systems rather than blaming individuals. Blame-heavy cultures drive problems underground, which is one of the most dangerous conditions for any QMS. When employees fear punishment, deviations go unreported, and small issues grow into major failures.

Leaders can build culture by recognizing good catches, closing the loop on employee suggestions, and treating investigations as learning opportunities.


9. Supplier and Outsourced Process Control

Few organizations make everything themselves. Raw materials, components, software, testing services, and contract manufacturing all come from outside, and quality problems at suppliers become the organization’s own problems.

A strong QMS manages external providers through:

  • Selection and qualification, based on capability, quality history, financial stability, and regulatory compliance.
  • Clear requirements, communicated through specifications, quality agreements, and purchase terms.
  • Ongoing monitoring, through incoming inspection, scorecards, audits, and performance reviews.
  • Corrective action processes to resolve supplier nonconformities.
  • Risk-based oversight, applying the most control to the most critical suppliers.

In regulated industries, responsibility cannot be outsourced. Manufacturers remain accountable for the quality of products made or tested on their behalf.


10. Operational Control: Delivering Consistent Results

Operational control is where the QMS meets daily work. Whether the output is a product or a service, the organization needs to plan and control delivery so that results are consistent.

Elements of strong operational control include:

  • Clear requirements for each product or service, agreed with the customer.
  • Design and development controls, including defined inputs, reviews, verification, validation, and change management.
  • Controlled conditions, such as qualified equipment, calibrated measuring instruments, suitable environments, and trained personnel.
  • In-process monitoring at critical points, rather than relying only on final inspection.
  • Identification and traceability, so that materials and outputs can be tracked when needed.
  • Handling of nonconforming output, including segregation, evaluation, and disposition so defective items do not reach customers.
  • Change control, ensuring that modifications to processes, materials, or systems are evaluated and approved before implementation.

The guiding idea is prevention. It is far cheaper to build quality into a process than to detect defects afterward and far cheaper still than to handle a recall or a lost customer.


11. Measuring Performance: Data, Audits, and Management Review

You cannot manage what you do not measure. A strong QMS uses data to see whether it is working and to drive decisions.

Key Performance Indicators

Useful quality metrics vary by organization but often include:

  • Customer satisfaction and complaint rates.
  • First-pass yield and defect rates.
  • On-time delivery.
  • Cost of poor quality (scrap, rework, returns, warranty).
  • Supplier performance.
  • Audit findings and closure times.
  • CAPA effectiveness and recurrence rates.

Metrics should be few enough to be understood, tied to objectives, and reviewed regularly. A dashboard full of numbers nobody acts on is worse than no dashboard.

Internal Audits

Internal audits, guided by ISO 19011, check whether the QMS conforms to requirements and is effectively implemented. Good audits go beyond box-ticking. They follow real processes, interview people, and examine evidence. Audit programs should be risk-based, giving more attention to high-risk or poorly performing areas, and auditors should be objective and trained.

Management Review

At planned intervals, top management reviews the QMS to confirm it remains suitable, adequate, and effective. Inputs include audit results, customer feedback, process performance, status of corrective actions, supplier performance, and changes in context or risk. The output should be concrete decisions about improvements, resource needs, and changes to the system.


12. Nonconformity, Corrective Action, and Continual Improvement

Problems are inevitable. What distinguishes a strong QMS is how it responds to them.

Correction deals with the immediate problem, such as quarantining a defective batch. Corrective action goes further by eliminating the cause to prevent recurrence. Preventive action, reflected in modern standards through risk-based thinking, addresses potential problems before they occur.

An effective CAPA process includes:

  1. Clear problem definition, with facts rather than assumptions.
  2. Containment, to protect customers and patients.
  3. Root cause analysis, using tools such as the 5 Whys, fishbone (Ishikawa) diagrams, or fault tree analysis.
  4. Action planning, with owners and due dates.
  5. Implementation and verification, confirming that the actions were completed.
  6. Effectiveness checks, confirming that the problem has not returned.

Weak CAPA systems are a leading cause of regulatory findings. Common failures include stopping at “operator error” as the root cause, closing actions without verifying results, and letting overdue actions pile up.

Continual improvement extends beyond fixing failures. The Plan-Do-Check-Act (PDCA) cycle provides a simple engine for it: plan a change, test it, check the results, and standardize what works. Lean, Six Sigma, and Kaizen methods offer additional structured approaches to eliminating waste and variation.


13. Technology and Digital Tools

Electronic quality management software can make a QMS more efficient and transparent. Modern platforms integrate document control, training management, deviation and CAPA tracking, audit management, supplier quality, and analytics in one place.

Benefits include faster workflows, automatic reminders, better traceability, and real-time visibility into quality performance. Advanced analytics can highlight trends that humans might miss, such as early signs of equipment drift or rising defect rates at a particular supplier.

However, technology does not fix a broken process. Digitizing a confusing procedure only makes the confusion faster. Organizations should simplify and standardize processes first, then choose tools that fit. In regulated settings, software used for quality records must also be validated and meet data integrity expectations.


14. A Practical Roadmap to Build a QMS

For organizations starting from scratch or rebuilding, a phased approach works best:

Phase 1: Commit and plan. Secure leadership commitment, appoint a quality lead, define scope, and identify applicable standards and regulations.

Phase 2: Assess the current state. Perform a gap analysis against the chosen standard to see what exists, what is missing, and what is not working.

Phase 3: Design the system. Map processes, define ownership, set the quality policy and objectives, and identify risks. Keep the design as simple as the organization allows.

Phase 4: Document and train. Develop the necessary procedures and records with input from the people who do the work, then train everyone on their responsibilities.

Phase 5: Implement. Run the processes, collect records, and refine procedures based on real-world experience.

Phase 6: Verify. Conduct internal audits and a management review to confirm the system works and to find weaknesses.

Phase 7: Certify or seek approval (if applicable). Undergo a third-party certification audit or regulatory inspection.

Phase 8: Sustain and improve. Keep monitoring, auditing, and improving. A QMS is never finished.

Expect the initial build to take several months for a small organization and longer for a large or highly regulated one. Rushing tends to produce paper systems that look good but are not used.


15. Common Pitfalls to Avoid

Even well-meaning organizations stumble on predictable problems:

  • Quality as a department, not a shared responsibility. When only the quality team cares about quality, the system becomes policing rather than prevention.
  • Over-documentation. Excessive procedures that no one reads or follows create a gap between paper and practice.
  • Copy-and-paste systems. Templates adopted without adaptation fail to reflect how work is actually done.
  • Audit-driven compliance. Systems that come alive only before an audit collapse in between.
  • Ignoring data. Collecting metrics without analyzing or acting on them wastes effort.
  • Superficial root cause analysis. Blaming individuals allows the underlying system flaw to persist.
  • Neglecting change management. Uncontrolled changes to processes, suppliers, or software are a frequent source of unexpected failures.
  • Lack of follow-through. Actions that are assigned but never verified make the system look healthy while problems grow.

Avoiding these comes down to honesty, simplicity, and discipline. The QMS should reflect reality, be as simple as possible, and be used every day.


Conclusion

A strong Quality Management System is not built by writing procedures alone. It grows from committed leadership, a clear understanding of customer and regulatory needs, well-defined processes, sensible risk management, competent and engaged people, reliable data, and a culture that treats problems as opportunities to learn. Each element supports the others. Processes without leadership drift, leadership without data guesses, and data without a culture of openness never tells the full story.

Organizations that invest in a real QMS gain more than certificates or clean inspection reports. They see fewer defects, less rework, more satisfied customers, stronger supplier relationships, and greater resilience when conditions change. Most importantly, they build the trust that every customer, patient, and regulator places in them.

Quality is not a destination but a continuing practice. The organizations that thrive are those that keep asking how they can do things better, and build systems that make the answer part of everyday work.


Guideline References

  1. ISO 9001:2015 – Quality management systems – Requirements.
  2. ISO 9000:2015 – Quality management systems – Fundamentals and vocabulary.
  3. ISO 9004:2018 – Quality management – Quality of an organization – Guidance to achieve sustained success.
  4. ISO 19011:2018 – Guidelines for auditing management systems.
  5. ISO 31000:2018 – Risk management – Guidelines.
  6. ISO 10002:2018 – Quality management – Customer satisfaction – Guidelines for complaints handling in organizations.
  7. ISO 13485:2016 – Medical devices – Quality management systems – Requirements for regulatory purposes.
  8. ICH Q9 (R1) – Quality Risk Management.
  9. ICH Q10 – Pharmaceutical Quality System.
  10. US FDA, 21 CFR Part 820 – Quality Management System Regulation (QMSR), incorporating ISO 13485:2016 by reference.
  11. US FDA, 21 CFR Parts 210 and 211 – Current Good Manufacturing Practice for Finished Pharmaceuticals.
  12. European Commission, EudraLex Volume 4, Part I, Chapter 1 – Pharmaceutical Quality System.
  13. WHO Technical Report Series No. 986, Annex 2 – WHO Good Manufacturing Practices for Pharmaceutical Products: Main Principles.

Building a Strong Quality Management System Read More »
Vendor Audit Questionnaire for Laminated Tube (Lamitube) Manufacturer

Vendor Audit Questionnaire for Laminated Tube: Below is a structured, regulator-aligned Vendor Audit Questionnaire tailored for a laminated tube (Lamitube) manufacturer supplying to pharmaceutical companies. The framework integrates expectations from World Health Organization, U.S. Food and Drug Administration, Pharmaceutical Inspection Co-operation Scheme, and European Medicines Agency guidelines (especially GMP for packaging materials, e.g., EU GMP Annex 1, WHO TRS 986/1019, FDA 21 CFR 210/211).


The questions are short, audit-friendly, and layered (Main → Sub-questions) so you can probe deeply during inspection.


🔷 1. Quality Management System (QMS)

Main Question:

How is the Quality Management System established, implemented, and maintained?

Breakdown:

  • Is there a documented Quality Manual aligned with GMP?
  • Are SOPs controlled, approved, and version-managed?
  • How is document lifecycle (issuance → archival) controlled?
  • Are quality objectives defined and periodically reviewed?
  • Is there a system for Quality Risk Management (ICH Q9 aligned)?
  • Are deviations, CAPA, and change control systems integrated?
  • Is management review conducted periodically?

🔷 2. Regulatory Compliance & Certifications

Main Question:

Does the company comply with applicable regulatory and GMP requirements?

Breakdown:

  • Which certifications are held (ISO 9001, ISO 15378, GMP)?
  • Are audits conducted by regulatory agencies or customers?
  • Are previous audit findings available with CAPA status?
  • Is there compliance with pharma packaging guidelines?
  • Are regulatory inspections documented and closed?

🔷 3. Personnel & Training

Main Question:

Are personnel qualified, trained, and compliant with GMP?

Breakdown:

  • Is there an organizational chart with defined responsibilities?
  • Are job descriptions documented?
  • How is initial and ongoing GMP training conducted?
  • Are training effectiveness evaluations performed?
  • Are personnel hygiene and gowning procedures defined?
  • Are operators trained in contamination control?

🔷 4. Premises & Facility Design

Main Question:

Is the facility designed to prevent contamination and mix-ups?

Breakdown:

  • Is material and personnel flow logically segregated?
  • Are clean/controlled areas defined (if applicable)?
  • Is there HVAC system qualification and monitoring?
  • Are pest control systems in place?
  • Are lighting, temperature, humidity controlled?
  • Are utilities (compressed air, water) monitored?

🔷 5. Equipment & Maintenance

Main Question:

Are equipment qualified, calibrated, and maintained?

Breakdown:

  • Is there a list of critical equipment?
  • Are IQ/OQ/PQ performed where applicable?
  • Is preventive maintenance planned and documented?
  • Are calibration records maintained?
  • Are breakdowns recorded and investigated?
  • Is cleaning of equipment validated or verified?

🔷 6. Raw Materials Control

Main Question:

How are raw materials (e.g., laminate, resins, inks) controlled?

Breakdown:

  • Are suppliers qualified and approved?
  • Are incoming materials tested against specifications?
  • Is there a quarantine → release system?
  • Are COAs verified?
  • Are storage conditions controlled?
  • Is FEFO/FIFO followed?

🔷 7. Production & Process Control

Main Question:

Are manufacturing processes controlled and validated?

Breakdown:

  • Are batch manufacturing records (BMR) available?
  • Are critical process parameters defined and monitored?
  • Is process validation performed?
  • Are line clearance procedures implemented?
  • Are in-process checks documented?
  • Is rejection/rework controlled?

🔷 8. Printing & Artwork Control (Critical for Lamitube)

Main Question:

How is printed artwork controlled to prevent errors?

Breakdown:

  • Is artwork approved by customers before printing?
  • Are master artworks securely controlled?
  • Are printing plates/cylinders controlled and verified?
  • Is line clearance done between different artworks?
  • Are barcode and text verification systems used?
  • Are reconciliation procedures in place?

🔷 9. Cleaning & Contamination Control

Main Question:

Are cleaning procedures effective to prevent contamination?

Breakdown:

  • Are cleaning SOPs defined for all areas?
  • Are cleaning records maintained?
  • Is cleaning validation required and performed?
  • Are residues (ink, adhesive) controlled?
  • Are cross-contamination risks assessed?

🔷 10. Laboratory Controls & Testing

Main Question:

Are QC laboratory operations reliable and compliant?

Breakdown:

  • Are specifications defined for tubes (dimension, leakage, print quality)?
  • Are test methods validated?
  • Is there an approved sampling plan?
  • Are instruments calibrated?
  • Are OOS (Out of Specification) handled properly?
  • Are retain samples stored?

🔷 11. Packaging & Finished Product Control

Main Question:

How are finished laminated tubes controlled and released?

Breakdown:

  • Are finished goods inspected before release?
  • Is batch traceability ensured?
  • Are packing materials controlled?
  • Are labeling and identification clear?
  • Is transport validation considered?

🔷 12. Storage & Distribution

Main Question:

Are storage and distribution conditions controlled?

Breakdown:

  • Are warehouses clean and organized?
  • Are environmental conditions monitored?
  • Are dispatch procedures documented?
  • Is traceability maintained during shipment?
  • Are damaged goods handled properly?

🔷 13. Deviations, CAPA & Change Control

Main Question:

Are quality issues effectively managed?

Breakdown:

  • Are deviations recorded and investigated?
  • Is root cause analysis performed?
  • Are CAPAs tracked for effectiveness?
  • Are changes evaluated for impact before implementation?
  • Is change control linked to validation?

🔷 14. Complaint Handling & Recall

Main Question:

Is there an effective system for complaints and recalls?

Breakdown:

  • Are customer complaints logged and investigated?
  • Is trending analysis performed?
  • Is there a mock recall system?
  • How quickly can products be traced and recalled?
  • Are regulatory authorities notified if required?

🔷 15. Data Integrity & Documentation

Main Question:

Is data integrity ensured across all systems?

Breakdown:

  • Are records attributable, legible, contemporaneous (ALCOA)?
  • Are electronic systems validated?
  • Are audit trails enabled?
  • Is access control implemented?
  • Are backups performed regularly?

🔷 16. Vendor & Supply Chain Control

Main Question:

How are upstream vendors controlled?

Breakdown:

  • Is there an approved vendor list (AVL)?
  • Are vendor audits conducted?
  • Are critical suppliers periodically re-evaluated?
  • Are agreements (Quality Agreements) in place?

🔷 17. EHS (Environment, Health & Safety)

Main Question:

Are safety and environmental controls implemented?

Breakdown:

  • Are MSDS available for chemicals (inks, solvents)?
  • Are hazardous materials stored properly?
  • Is waste disposal compliant?
  • Are fire safety systems installed?
  • Are emergency procedures defined?

🔷 18. Continuous Improvement

Main Question:

Does the company focus on continuous improvement?

Breakdown:

  • Are KPIs defined (rejection rate, complaints)?
  • Are internal audits conducted?
  • Are improvement projects documented?
  • Is trend analysis performed?

✅ Pro Tips for Strong Vendor Audit (Practical Execution)

  • Focus heavily on:
    • Artwork control (highest risk in tubes)
    • Traceability (batch → raw material → dispatch)
    • Line clearance & mix-up prevention
  • Ask for live records, not just SOPs.
  • Perform shop-floor verification (not just QA office review).
  • Cross-check:
    • BMR vs actual production
    • COA vs test raw data
  • Look for data integrity red flags (backdating, overwriting, missing logs)


🔴 SERIES–2: ADVANCED / FORENSIC VENDOR AUDIT QUESTIONS


🔷 1. QMS Effectiveness (Beyond Documentation)

Main Question:

How do you demonstrate that your QMS is effective, not just compliant?

Breakdown:

  • Show last 12 months trend of deviations vs CAPA closure time
  • What % of CAPAs are delayed? Justify
  • Provide example where CAPA failed → what was done next?
  • How do you ensure recurrence does NOT happen?
  • Are KPIs linked to management incentives?
  • Show evidence of risk-based decision making, not SOP-driven only

🔷 2. Data Integrity (Critical – High Risk Area)

Main Question:

How do you ensure ALCOA+ compliance in real operations?

Breakdown:

  • Show raw data vs reported COA → any mismatch?
  • Are there any blank spaces, overwriting, backdating in records?
  • Who reviews audit trails and how frequently?
  • Can operators delete or modify data?
  • Show one real audit trail for a batch
  • What controls prevent “testing into compliance”?
  • How do you detect unofficial records (shadow documentation)?

🔷 3. Deviation Investigation Depth

Main Question:

How robust is your root cause investigation?

Breakdown:

  • Show a major deviation report (last 6 months)
  • Was root cause scientifically justified or assumed?
  • Was 5 Why / Fishbone / FMEA used?
  • Were multiple root causes considered?
  • Was QA independent in conclusion approval?
  • How do you verify root cause correctness?

🔷 4. Change Control – Hidden Risks

Main Question:

How do you ensure changes do not introduce unseen risks?

Breakdown:

  • Show a recent change control affecting production
  • Was risk assessment formal (ICH Q9)?
  • Were validation/revalidation requirements evaluated?
  • Was customer notified (for critical changes)?
  • Were changes implemented before approval (backdoor change)?
  • How do you track temporary changes?

🔷 5. Process Validation – Real Assurance

Main Question:

How do you prove your process is consistently capable?

Breakdown:

  • Show Process Validation Report
  • Are worst-case conditions included?
  • What is the process capability (Cp/Cpk)?
  • How are critical parameters justified?
  • Is continued process verification (CPV) performed?
  • Show trend of in-process rejection over time

🔷 6. Artwork & Printing – Zero Error Expectation Area

Main Question:

How do you ensure zero artwork error, considering high regulatory risk?

Breakdown:

  • Show real case of artwork error → what happened?
  • Is there 100% vision inspection or manual?
  • How is mix-up between similar artworks prevented?
  • Are cylinders uniquely coded and verified?
  • Is there dual verification before printing?
  • What is reconciliation tolerance for printed tubes?

🔷 7. Line Clearance – Practical Effectiveness

Main Question:

How do you guarantee zero mix-up between batches/products?

Breakdown:

  • Demonstrate a live line clearance
  • Who verifies clearance (production vs QA)?
  • Are previous batch remnants physically checked?
  • Are labels/artwork fragments controlled?
  • Is photographic evidence used?

🔷 8. Supplier Risk Management

Main Question:

How do you control risks from your raw material suppliers?

Breakdown:

  • Show supplier risk categorization
  • How often are critical suppliers audited?
  • Any supplier disqualified recently? Why?
  • How do you verify supplier COA reliability?
  • Do you perform skip testing or full testing?

🔷 9. Laboratory Control – Reliability Check

Main Question:

How do you ensure QC results are scientifically reliable?

Breakdown:

  • Show OOS investigation (real case)
  • Was Phase I / Phase II investigation followed?
  • Are analysts trained on data integrity?
  • Any retesting without justification?
  • How are reference standards controlled?

🔷 10. Rejection, Rework & Scrap Control

Main Question:

How do you prevent rejected materials from re-entering the system?

Breakdown:

  • Show rejected batch records
  • Is rework scientifically justified and approved?
  • How is scrap physically destroyed?
  • Any case of unauthorized reprocessing?
  • Are rejection trends analyzed?

🔷 11. Traceability (Backward & Forward)

Main Question:

Can you trace any product within minutes?

Breakdown:

  • Perform mock traceability test:
    • Finished tube → raw material batch
    • Raw material → all affected customers
  • Time required to retrieve data?
  • Is system manual or ERP-based?
  • Any gaps in traceability observed?

🔷 12. Complaint Handling – True Root Cause

Main Question:

How do you ensure complaints lead to real improvement?

Breakdown:

  • Show last 5 complaints
  • Are trends analyzed (e.g., print defect, leakage)?
  • Were CAPAs effective?
  • Any repeated complaints for same issue?
  • Was customer feedback incorporated?

🔷 13. Recall System – Practical Readiness

Main Question:

How prepared are you for a real recall?

Breakdown:

  • Show last mock recall report
  • Time taken to complete recall simulation?
  • % traceability achieved?
  • Are customers informed in simulation?
  • Are regulatory requirements considered?

🔷 14. Cleaning Validation (Critical for Cross-Contamination)

Main Question:

How do you ensure no carryover of ink/chemical residues?

Breakdown:

  • Are worst-case products selected?
  • What is acceptance criteria (scientific or arbitrary)?
  • Are swab/recovery studies performed?
  • Is visual cleanliness justified scientifically?
  • Any failure observed in cleaning validation?

🔷 15. Environmental Control (Especially Printing Area)

Main Question:

How do environmental factors impact product quality?

Breakdown:

  • Are temp/humidity linked to print quality?
  • Any environmental excursions recorded?
  • Impact assessment done?
  • Are sensors calibrated?

🔷 16. Warehouse – Hidden GMP Gaps

Main Question:

How do you prevent mix-up and deterioration in storage?

Breakdown:

  • Are status labels (Quarantine/Released/Rejected) clear?
  • Any case of wrong material issuance?
  • Are returns segregated?
  • Is pest control effective (show logs)?

🔷 17. Internal Audit Effectiveness

Main Question:

Are internal audits truly identifying gaps?

Breakdown:

  • Show last internal audit report
  • Were critical findings raised?
  • Any repeat observations?
  • Is QA independent from production?

🔷 18. Management Commitment (Reality Check)

Main Question:

How does top management ensure GMP compliance?

Breakdown:

  • Are resources sufficient (manpower, budget)?
  • Any production vs quality conflict?
  • Example where batch was rejected despite pressure
  • Are quality metrics reviewed at top level?

🔥 HIGH-IMPACT AUDIT TECHNIQUES (Use During Audit)

  • Ask: “Show me now” instead of “Do you have”
  • Cross-question same topic with:
    • Operator
    • Supervisor
    • QA
  • Pick random batch → trace everything
  • Compare:
    • SOP vs actual practice
  • Watch for:
    • Delayed entries
    • Too-perfect records
    • Identical handwriting


🔴 SERIES–3: TRAP QUESTIONS (HIGH-IMPACT AUDIT TOOL)

⚠️ How to use:

  • Ask indirectly
  • Cross-check answers between departments
  • Always demand live evidence

🔷 1. “Show Me Now” Trap (Reality vs Documentation)

Main Trap Question:

“Can you show me a batch currently under processing and all associated live records?”

Trap Breakdown:

  • Does the operator hesitate or call QA first?
  • Are entries made in real-time or backfilled?
  • Do timestamps align with actual process time?
  • Is handwriting consistent or pre-filled?

👉 Red Flag: Records updated just before showing


🔷 2. Data Integrity Trap (Backdating Detection)

Main Trap Question:

“Pick any record from last week—can you explain when exactly this entry was made?”

Trap Breakdown:

  • Compare:
    • Ink color / pen variation
    • Time gaps between entries
  • Ask operator:
    • “Did you write this immediately or later?”

👉 Red Flag: Same pen, same handwriting for full shift data


🔷 3. Shadow Documentation Trap

Main Trap Question:

“Do operators ever use rough sheets before final entry?”

Trap Breakdown:

  • Check drawers, behind machines
  • Ask casually: “How do you remember readings before writing?”

👉 Red Flag: Unofficial notebooks / loose papers


🔷 4. Deviation Suppression Trap

Main Trap Question:

“In the last 3 months, did any process fail but was not recorded as deviation?”

Trap Breakdown:

  • Cross-check:
    • Maintenance logs
    • QC OOS
    • Production downtime
  • Compare with deviation register

👉 Red Flag: Events exist but no deviation raised


🔷 5. CAPA Effectiveness Trap

Main Trap Question:

“Show a deviation that happened twice—why did CAPA fail first time?”

Trap Breakdown:

  • Check recurrence
  • Evaluate depth of root cause

👉 Red Flag: Same issue repeating with different wording


🔷 6. Artwork Mix-up Trap (Critical for Tubes)

Main Trap Question:

“What prevents two similar artworks from being mixed during printing?”

Trap Breakdown:

  • Physically verify:
    • Cylinders
    • Printed tubes
  • Ask operator to explain difference

👉 Red Flag: Reliance only on visual check without system control


🔷 7. Line Clearance Trap (False Compliance)

Main Trap Question:

“Can we check the previous product run on this line?”

Trap Breakdown:

  • Look for:
    • Leftover labels
    • Tubes under machine
  • Ask: “Who verified clearance?”

👉 Red Flag: Clearance signed but physical evidence remains


🔷 8. Training Effectiveness Trap

Main Trap Question:

Ask operator directly:
“Explain what you will do if you find a defective tube?”

Trap Breakdown:

  • Compare with SOP
  • Check confidence level

👉 Red Flag: SOP exists but operator unaware


🔷 9. QC Testing Manipulation Trap

Main Trap Question:

“Have you ever repeated a test because the first result was not acceptable?”

Trap Breakdown:

  • Ask follow-up:
    • “Was it documented?”
  • Check raw data vs final COA

👉 Red Flag: Retesting without investigation


🔷 10. OOS Handling Trap

Main Trap Question:

“Show me an OOS result and explain full investigation”

Trap Breakdown:

  • Check:
    • Hypothesis-driven investigation
    • Lab error justification

👉 Red Flag: Blaming analyst without proof


🔷 11. Change Control Bypass Trap

Main Trap Question:

“Have you ever implemented a change urgently before approval?”

Trap Breakdown:

  • Cross-check with production/engineering
  • Verify temporary changes

👉 Red Flag: Verbal approvals / undocumented changes


🔷 12. Supplier COA Trust Trap

Main Trap Question:

“Do you always test raw materials or rely on supplier COA?”

Trap Breakdown:

  • Ask:
    • “When was last full testing done?”

👉 Red Flag: Blind trust in supplier without verification


🔷 13. Traceability Stress Test (Live Challenge)

Main Trap Question:

“Trace this finished tube to raw materials within 10 minutes”

Trap Breakdown:

  • Observe:
    • Time taken
    • Data gaps

👉 Red Flag: Delays, incomplete traceability


🔷 14. Scrap Control Trap

Main Trap Question:

“What happens to rejected printed tubes?”

Trap Breakdown:

  • Physically inspect scrap area
  • Check destruction records

👉 Red Flag: Scrap accessible / reusable


🔷 15. Warehouse Mix-up Trap

Main Trap Question:

“Can you show any wrongly placed material found in last audit?”

Trap Breakdown:

  • Check labeling system
  • Verify segregation

👉 Red Flag: Same-looking materials stored together


🔷 16. Environmental Excursion Trap

Main Trap Question:

“What happens if humidity exceeds limit during printing?”

Trap Breakdown:

  • Ask for real case
  • Check deviation link

👉 Red Flag: No impact assessment


🔷 17. Internal Audit Integrity Trap

Main Trap Question:

“Why were no critical observations found in last internal audit?”

Trap Breakdown:

  • Compare with your findings

👉 Red Flag: Internal audit too “clean”


🔷 18. Management Pressure Trap

Main Trap Question:

“Has production ever continued despite quality concern?”

Trap Breakdown:

  • Ask different levels separately

👉 Red Flag: Conflicting answers


🔷 19. “Too Perfect System” Trap

Main Trap Question:

“You have zero deviations last month—how is that possible?”

👉 Red Flag: Indicates underreporting


🔷 20. Contradiction Trap (Powerful Technique)

Method:

Ask same question to:

  • Operator
  • QA
  • Supervisor

👉 Compare answers

Example:

  • “Who performs line clearance?”
  • “Who approves deviation?”

👉 Red Flag: Different answers


🔥 CRITICAL OBSERVATION SKILLS (MOST IMPORTANT)

During audit, continuously watch for:

  • ⛔ Entries written in same handwriting
  • ⛔ No corrections in months (unrealistic)
  • ⛔ Fearful operators
  • ⛔ QA answering everything (production silent)
  • ⛔ Delayed data retrieval
  • ⛔ “We always follow SOP” without explanation

🧠 PRO AUDITOR STRATEGY (FIELD-LEVEL)

  • Never rely on verbal answers
  • Always:
    • See → Verify → Cross-check
  • Use silence pressure (wait after asking)
  • Randomly select:
    • Batch
    • Operator
    • Record

✅ If you want FINAL LEVEL (ELITE)

I can prepare:

🔷 “Audit Scoring System”

  • Critical / Major / Minor classification
  • Risk-based scoring (FDA-style)

🔷 “Lamitube-Specific Risk Matrix”

  • Leakage risk
  • Print toxicity risk
  • Extractables/leachables concern

🔷 “DGDA Bangladesh Audit Angle”

  • Local regulatory expectations
  • Common inspection findings in Bangladesh

Vendor Audit Questionnaire for Laminated Tube (Lamitube) Manufacturer Read More »

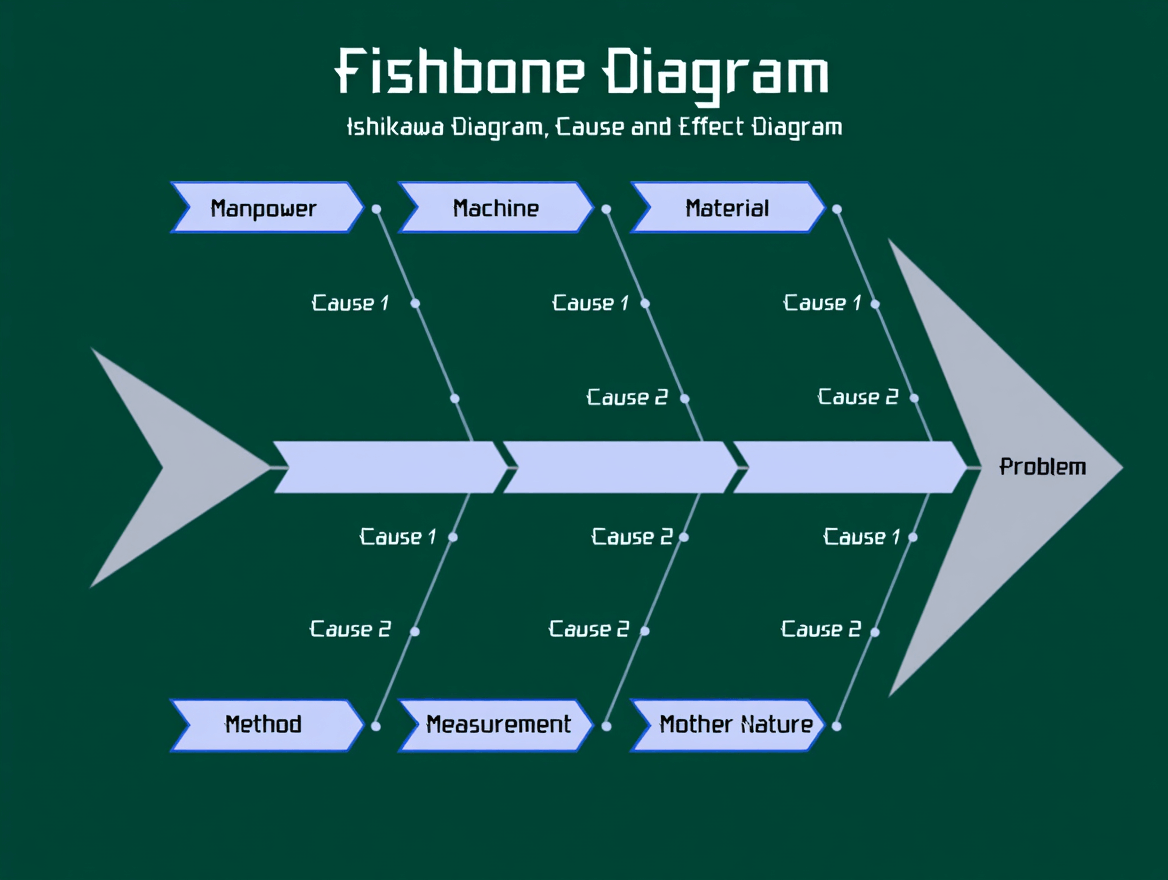
What is a Fishbone Diagram, how is it used?

A Fishbone Diagram (also called Ishikawa Diagram or Cause-and-Effect Diagram) is a visual tool used to systematically identify and organize all possible causes of a specific problem or effect. It was developed by Dr. Kaoru Ishikawa in the 1960s.

The diagram looks like a fish skeleton:

  • The “head” of the fish is the problem/effect you want to solve.
  • The “bones” branching off the spine are the major categories of causes.
  • Smaller bones branching off are the specific causes within each category.

Why use it?

  • Forces teams to think through all possible causes (not just the obvious ones)
  • Prevents jumping to conclusions
  • Encourages cross-functional collaboration
  • Very useful in root cause analysis (RCA), quality improvement, Six Sigma, Lean, etc.

How to Create and Use a Fishbone Diagram (Step-by-Step)

  1. Define the problem clearly
    Write it in a box on the right (the “head”). Be specific.
    Example: “Customer complaints about late deliveries increased by 40% in Q3”
  2. Draw the main spine
    A horizontal arrow pointing to the problem.
  3. Identify major cause categories (the big bones)
    Common categories (the 6 Ms for manufacturing, or adapt to your industry):

    • Man (People)
    • Method (Processes)
    • Machine (Equipment/Technology)
    • Material
    • Measurement
    • Mother Nature (Environment)

    For service/industry, people often use:

    • People, Policies, Procedures, Plant/Technology, etc.
  4. Brainstorm all possible causes
    Ask “Why does this happen?” repeatedly (5 Whys technique helps).
    Write each cause as a branch off the relevant category.
  5. Go deeper
    For each cause, ask “Why?” again and add sub-causes (smaller bones).
  6. Analyze and prioritize
    Circle the most likely root causes (use voting, data, Pareto, etc.).
  7. Develop action plan
    Address the confirmed root causes.

Practical Case Study: Restaurant Getting Complaints About Cold Food

Problem (Head of the fish):
“Customers frequently complain that food arrives cold” (complaints rose from 3 to 18 per week)

Fishbone Diagram Categories Used:
We used 6 categories suitable for a restaurant:

  1. People (Staff)
    • Servers forget to check food temperature before leaving kitchen
    • New staff not trained on urgency
    • Kitchen and serving staff not communicating
    • Too few servers during peak hours → delay
  2. Processes (Methods)
    • No standard procedure for checking food temperature
    • Food waits too long on the pass before being served
    • Tickets not prioritized correctly during rush
    • No “food ready” notification system between kitchen and floor
  3. Equipment (Machines)
    • Heat lamps broken or insufficient
    • Plates not pre-heated
    • Delivery trays too small → food stacked and cools faster
    • Old warming drawers not maintaining temperature
  4. Materials
    • Some dishes (e.g., pasta) cool faster than others
    • Large portion sizes take longer to eat → perceived as cold
  5. Environment (Mother Nature/Place)
    • Dining room AC set too low near some tables
    • Long walking distance from kitchen to farthest tables
    • Draft from entrance door
  6. Measurement (or Management)
    • No tracking of time from “food ready” to “delivered to table”
    • No temperature checks logged
    • No customer feedback analyzed by dish/table location

Root Causes Identified After Investigation (circled on the diagram):

  • Heat lamps were broken for 3 weeks (maintenance backlog)
  • No policy to pre-heat plates
  • Average time from kitchen to table was 4.5 minutes during peak (target should be <2 min)
  • Servers overloaded: 1 server handling 9 tables instead of max 6

Actions Taken:

  1. Repaired/replaced heat lamps same week
  2. Implemented plate pre-heating in salamander broiler
  3. Added a runner position during peak hours
  4. Created a simple checklist: food temperature >60°C before leaving pass
  5. Installed digital ticket system with timers

Result:
Cold food complaints dropped from 18 to 2 per week within one month.

What is a Fishbone Diagram, how is it used? Read More »

Data Integrity and Why Is It So Important?

Data Integrity can be defined as it is critical part to the plan, execution & practice of system that processes, stores & or retrieves data and maintenance & assurance of data accuracy and consistency throughout its life cycle.

Another best alternative term is Data quality which is sometimes defined as the proxy term of data integrity. It is the wide term & it can be used in different meaning depending on the framework. For data integrity, data validation is prerequisite. Data corruption is the just opposite term of Data integrity.

Also Read

What is the difference between Data and Information?

Confirm that data is recorded exactly as projected and most of the cases Data Integrity technique is the same for all type of probable data source.
The main target regarding is to prevent any type of alteration of the exact data recorded during collection of data and it can’t change at any cost. At the time of retrieval of data, it must be same as collected before.

Data Integrity denote the accuracy & consistency of data over its lifecycle. Sensitive data may loss upon uses of Negotiated data. Under this consideration, continuing it is a main focus of many enterprise security solutions. During replicated or transferred of data, it shall be intact and unaltered. To ensure the integrity of data, Error checking methods and validation procedures are typically best way to serve the same.

Don’t confuse with Data Integrity and Data security, it is process to prevent unauthorized entry to the projected data or protecting data from unauthorized expert.
Any unintended changes to data as the consequence of a storage, retrieval or processing operation, including malicious intent, unexpected hardware failure, and human error, is failure of data integrity.

Failure of Data Integrity means any unplanned changes to data as the consequence of storage, unexpected hardware failure, retrieval or human error or processing operation malicious intent etc. If you unable to protect your data from unauthorized change then it can define as data security failure.

Data Integrity denote that it is a state as well as process, so a confusion may arise. Data Integrity is a state as it denotes a data set is both valid and accurate. Validation methods & Error checking are considered as Data Integrity processes.

Study of Data Integrity

Data Integrity is significant for several reasons & need to maintain the same. Data Integrity confirms searchability, recoverability, connectivity, & traceability. To increases stability & performance as well as improving reusability & maintainability; need to protect validity & accuracy of data. Data plays a major role to drive enterprise decision-making and data undergo several stages of changes to form raw to format to become more practical and identifying relation between them. In modern enterprises Data Integrity consider the top most priority.

In a database system, Data Integrity may be Compromised in various type of ways. In the following ways, Data Integrity may be compromised-
[][]Compromised hardware, like device or disk crash
[][]Compromise of Physical devices
[][]Cyber threats, Bugs, Viruses/Malware, Hacking, & other unfamiliar process
[][]Human error, whether unintentional or malicious

Transfer errors, unintended alterations or data compromise during transfer from one device to another device.
Most of the cases, some type of data security may protect this data compromises. Data duplication is the critical para meter for data security as well as data backup process.

Data Integrity & Databases

Data Integrity encompasses strategies for data specifying, retention, or guaranteeing the length of time data. Any lessening of enforcement could cause errors in the data; all of the rules shall be consistently & regularly implemented to all data entering the system.

At time of data input, checking system shall be implement which will lessen up the number of data error for the system. Data Integrity rules shall be strictly implemented to the system which will save troubleshooting time, erroneous data subsequently errors to algorithms.

A standard Data Integrity rules must have the strict definition regarding data relation; as which type data shall link with which type of data. A selling record of a item of certain product may be linked with the specific product but is shall not be related to unrelated data such as company asset, policy, loan, promotion etc. Based on predefined rules, it may be included check & correction system for the invalid a data.

Data derivation rules shall be applicable, mentioning data derivation procedure that how a data value shall be derived based on contributors, conditions and algorithm of the system. Re-derived procedure for data value shall be mention on which condition shall be considered for this process.

Types of Data Integrity

Organizations can maintain Data Integrity through integrity constraints, which define the rules and procedures around actions like deletion, insertion, and update of information. The definition of Data Integrity can be enforced in both hierarchical and relational databases, such as enterprise resource planning (ERP), customer relationship management (CRM), and supply chain management (CRM) systems.

Organizations can achieve it through the following:

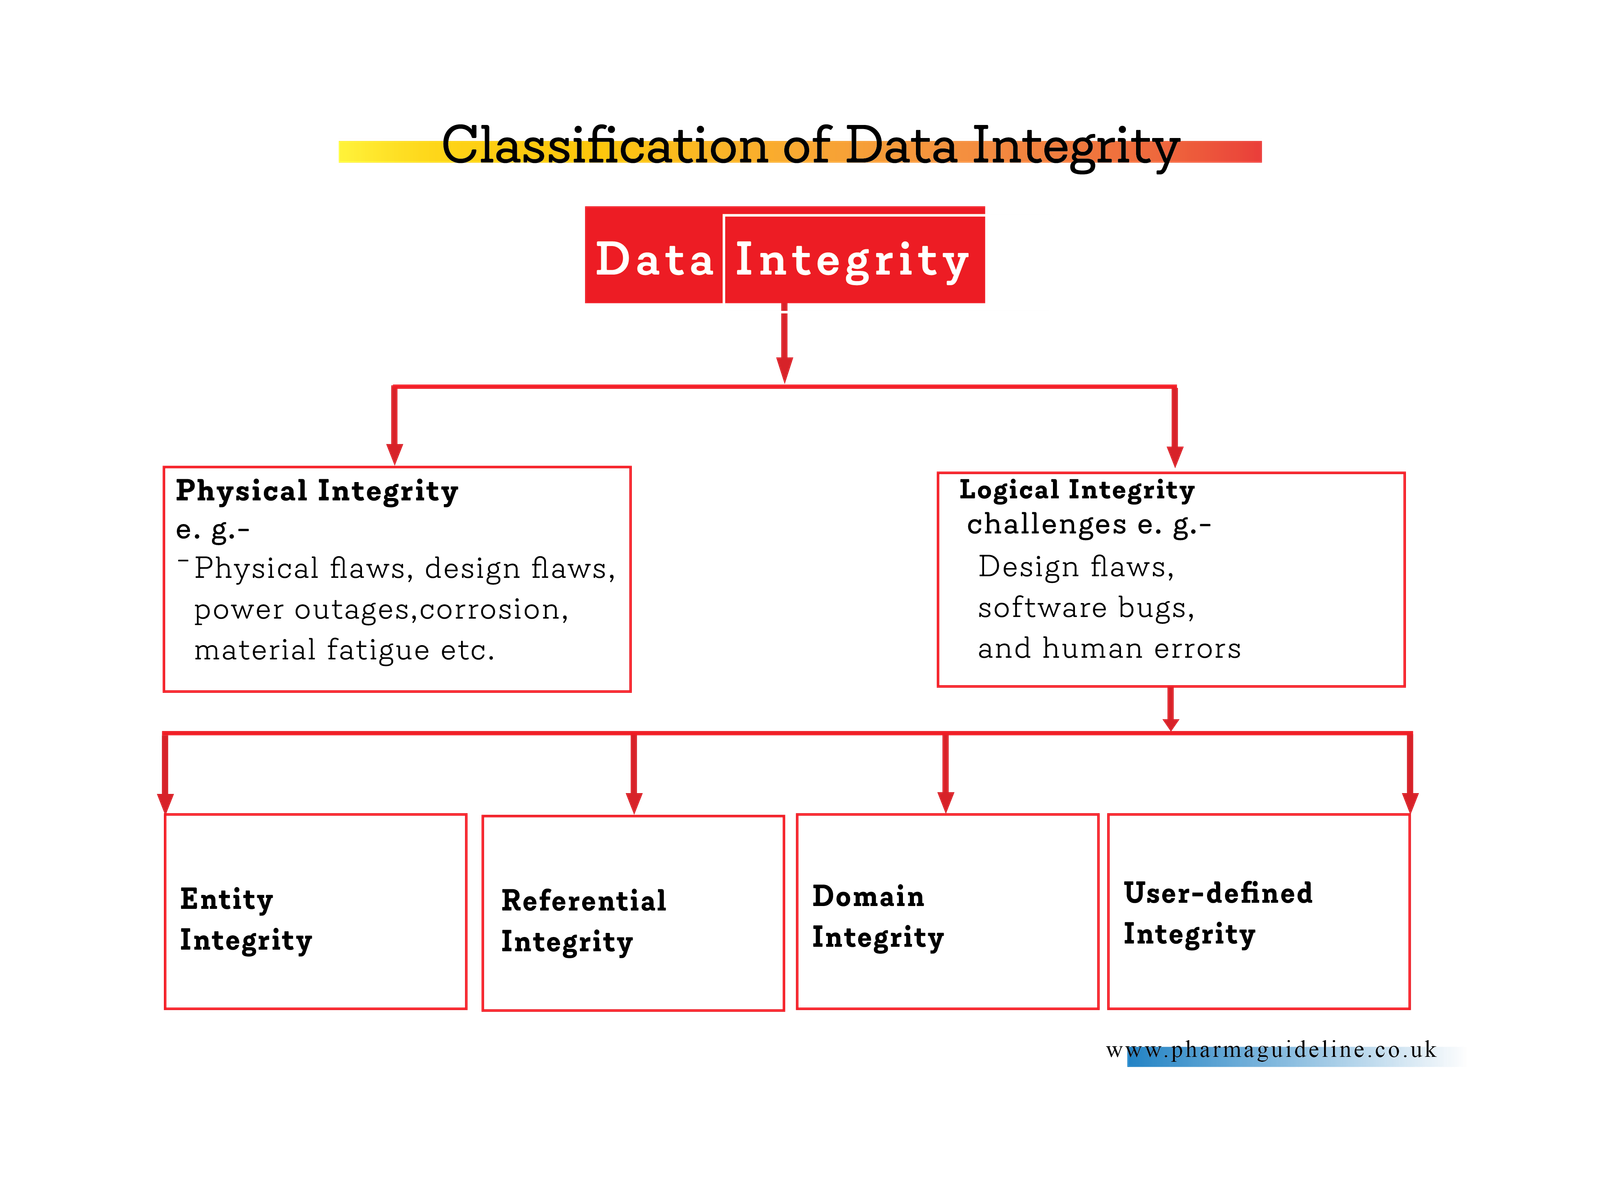
Physical Integrity

Physical integrity deals with challenges which are associated with correctly storing and fetching the data itself.

Physical integrity indicates the right storing & fetching the data itself & its associated series of challenges. Various types of Challenges are involved with it such as Physical flaws, design flaws, power outages, electromechanical faults, corrosion, material fatigue, natural disasters, environmental hazards such as ionizing radiation, high temperatures, pressures & g-forces [the force of gravity].

Various methods are available to maintain the physical integrity such as UPS [uninterruptible power supply], redundant hardware, various type of RAID arrays, error-correcting memory, radiation hardened chips, clustered file system, watchdog timer & cryptographic hash function for critical system.

Error-correcting codes is extensively use as error detecting algorithms in Physical integrity of data management systems. Simpler checks & algorithms as the Damm algorithm or Luhn algorithm is use to detect Data Integrity errors. This system is use to uphold Data Integrity at manual transcription from one computer system to another computer system through a human intermediary such as credit card numbers. Hash functions are more beneficial to detect Computer-induced transcription errors.

These techniques are used together to ensure various degrees of Data Integrity in production systems such as a fault-tolerant RAID array may be use to configured a computer system but in silent data corruption block-level checksums might not provide.

In a nutshell, Physical integrity means protecting the accuracy, correctness, and wholeness of data when it is stored and retrieved. This is typically compromised by issues like power outages, storage erosion, hackers targeting database functions, and natural disasters, which prevent accurate data storage and retrieval.

Logical Integrity

It is concerned with correctness or rationality of a piece of data provide a particular context. It denotes the topic such as Entity integrity & referential integrity in a relational database system. Design flaws, software bugs, and human errors are the major challenges. Foreign key constraints, check constraints, program assertions, and other run-time sanity checks are the common methods to ensure logical integrity.

Design flaws & Human errors, both are the common problems for physical and logical integrity which must be properly deal with the contemporaneous requests to record and retrieve data. Physical error for a specific data system is more critical than logical error. If a data system suspected to logical error, it can be reused by overwriting with the new one but if it faces physical error then the data sector is totally used of its own condition.

Logical integrity ensures that data remains unchanged while being used in different ways through relational databases. This approach also aims to protect data from hacking or human error issues but does so differently than physical integrity.

Logical integrity comes in four different formats:

Entity Integrity

Entity integrity is a feature of relation systems that store data within tables, which can be used and linked in various ways. It relies on primary keys and unique values being created to identify a piece of data. This ensures data cannot be listed multiple times, and fields in a table cannot be null.

Referential Integrity

Referential integrity is a series of processes that ensure data remains stored and used in a uniform manner. Database structures are embedded with rules that define how foreign keys are used, which ensures only appropriate data deletion, changes, and amendments can be made. This can prevent data duplication and guarantee data accuracy.

Domain Integrity

Domain integrity is a series of processes that guarantee the accuracy of pieces of data within a domain. A domain is classified by a set of values that a table’s columns are allowed to contain, along with constraints and measures that limit the amount, format, and type of data that can be entered.

User-defined Integrity

User-defined integrity means that rules and constraints around data are created by users to align with their specific requirements. This is usually used when other integrity processes will not safeguard an organization’s data, allowing for the creation of rules that incorporate an organization’s Data Integrity measures.

Types of Integrity Constraints

A set of integrity constraints or rules are followed to implement Data Integrity in a database system. Relational data model suggests the three types of integrity constraints such as domain integrity, entity integrity, & referential integrity.

First of all, Entity integrity denote the concept of a primary key. As per this system, Entity integrity states that every table shall contain a primary key and existing column or columns of the table shall be identified by the primary key & it shall be inimitable and not null.

Concept of a foreign key denote Referential integrity. As per referential integrity rule, any foreign-key value can only be in one of two states. In general condition, foreign-key value refers to a primary key value of some table in the database system. Sporadically, a foreign-key value can be null and this will rest on on the rules of data owner. Under these circumstances, it can be stated here that this relationship is unknown or there is no relationship between the objects represented in the database system.

All columns in a relational database must be stated upon a defined domain is the main concern of the Domain integrity. In the relational data model, the primary unit of data is the data item. This type of data items is known as atomic or non-decomposable. A set of values of the same type is defined as domain. Actual values appearing in the columns of a table are drawn from the Domains which are considered as pools of values.

As the User-defined integrity are set by the specific user with a set of rules which is not related to domain, entity, and referential integrity classes. The database which supports these features, it is the sole responsibility of the database to confirm Data Integrity and reliability model for the data storage & retrieval system. The database which does not support these features then it is the accountability of the applications to confirm Data Integrity though the database supports the consistency model for data storage & recovery process.

A well-controlled single and well managed data-integrity system increases.

[][]Maintainability
=>All Data Integrity administration commences from a single centralized system.

[][]Performance
=>As a single operation unit, all Data Integrity operations perform as consistency model.
[][]Re-usability
=>A single centralized Data Integrity system provides the all applications benefit.
[][]Stability
=>As avoid the multiple system, so Data Integrity operation performs sound as well as better retrieval on a one centralized system.

Examples
Data-integrity mechanism is often considered as the parent-and-child connection of interrelated records. When one or more related child records exist for a parent record then all of referential integrity methods are handled by database itself & inevitably ensures accuracy & integrity of data, so no child record can exist without parent record and subsequently no parent drops their child records. In this system if the parent record owns any child records, then no parent record can be deleted and all of the process handled by the database system.

File systems

File system including Ext, JFS, UFS, XFS, and NTFS or hardware RAID solutions can’t provide satisfactory protection against Data Integrity problems. Some special file system such as BTRFS and ZFS use for silent data corruption can provide extra protection for data integrity. Upon provide this protection and being chance raise to corrupt the data then such file system can construct the data is widely known as end-to-end data protection.

Data Integrity as applied to various industries
[][]FDA has created the draft guidance for Data Integrity system as per 21 CFR Parts 210–212.[12] for pharmaceutical manufacturers. Same guideline has been developed by UK [2015], Switzerland [2016], and Australia [2017].
[][]Data Integrity also addressed by ISO as per ISO 13485, ISO 14155, and ISO 5840.
[][]FINRA [Financial Industry Regulatory Authority], implement the Data Integrity system on 2018 under technology change management policies and procedures” and Treasury securities reviews as Data Integrity problem found in 2017 on automated trading and money movement surveillance systems.

Why Data Integrity Matter?

Now a days Data are becoming more available, a smart business strategy which are using to make decision are obtaining the several times benefits.
According to recent research, a data driven organization is more than 23 times better performer in customer acquisition, nine times more performer to retain their customer and more than 19 times profitable to their competitor.

As the power of data is increasing day by day, so Data Integrity shall be valued properly and its importance can’t be denied at current situation. Presence of any type of error in data can spoil the total organization goal. A data driven organization must protect their database system at any cost to provide better security solution.

Threats involve in Data Integrity

[][]Human error:
It may arise in such case of transferring of data manually from one share drive to another, copying data from one spreadsheet to another and subsequently deleting of row or column of a spreadsheet. Storing data on excel sheet may cause formatting problem during manually data transferring process. Updating of excel sheet from old version to new one may cause formatting problem of subsequent data.

[][]Inappropriate format:
Stored data on Microsoft excel based on cell referencing may not accurate in different format. Failure to determine the same may case Data Integrity problem.

[][]Collection error:
During collecting any type of data, proper precaution shall be taken. Collecting of data on wrong method may cause storing of incomplete data and actual data may not represent the total situation.

[][]Internal security breaches:
If the database system hacked by third part or internal or external competitor may cause serious Data Integrity failure.

Why is Data Integrity Important?

Generally, a specific individual or group of people are involved in database system of a organization. Problem arises when multiple people are responsible to operate the database system. Anyone of the team member may not aware about Data Integrity of the organization, then all of the individual shall educate regarding protection of the database system and tech them the importance of data quality, accuracy, completeness etc. & they must learn how to combat when potential data security threat arises.

If all of the team member are aware about the Data Integrity and know its importance then it is very much effect to maintain the database system.

A better Data Integrity system can save company effort, time as well as valuable overhead cost. Wrong decision may take place based on inappropriate data. Data driven organization always take critical decision based on available data, if Data Integrity of that organization are compromised by any situation, then inappropriate result may arise and the organization suffer in long run.

Data always help to make important decision additionally it protect your company image. If you are collecting your customer information then protect the collect data which you have collect from your individual customer, failure to protect the customer data in proper database system may leak your customer information to another one cause image problem of the company and also mispresent your customer to other party.

Any type of customer information may be tracked and may be asked or run a campaign over them to collect specific target data. All type of collected information may not be sensitive as SSN [Social Security number]. To protect your valuable customer, you have to take a proper step regarding Data Integrity of your existing system.

a company staff always demand to data access to the database to trace any type of data on time manner upon request. He needs to uninterrupted access to the data system. For this reason, Data Integrity is so important for the organization. Data Integrity confirm the traceability & searchability of the data from its mother source.

Effective data accuracy and data protection shall be confirmed to increase Data performance and its stability. It is very much crucial to confirm the completeness & integrity of data. Compromised data always carry the wrong value for the organization & is of no use for most companies.

Same scenario arises for big data management. It is very much important to secure the big data management system as well as to maintain the total database system. All type data is totally worthless Without integrity & accuracy. Your data can be compromised If you fail to do the same. Under this circumstances, awkward & expensive data audit trails shall be mandatory to find out error & recover the total database system.

Most companies have set specific goals for their data & it is now more important than ever. But if integrity is not assured then data is not of much use. If data loss, corrupted or compromised then data can considerably damage any type of business. To maintain it, data security shall be confirmed using proper tools.

Compromised Data is the big challenge to maintain data integrity. There are several ways to compromised the valuable data. Todays almost all of the data are digital and store then in the same than traditional method and its transferred vary rapidly in different places of the globe. So, security shall be considering first & also the collection data is main concert maintain its integrity.

Data can be unaltered if data transferring occur maintaining valid system. Every time Data is moving from one place to another and it is not static, every user of the specific system is using the data and transferring the same in different way.

Management of Data Integrity

A group of steps are available where you can maintain & achieve better Data Integrity for your organization.

Collection of Accurate, Complete, and High-Quality Data

Quality of data depends on the collecting process of the projected data; a collection process is crucial and collection depends on the proper collection method. Failure to select proper data collection technic may cause collection of erroneous data. Sourcing is prerequisite to collect data. Ensure the high-quality data source may provide one step forwards to competitor.

Meticulously Check for Errors

To make common error during collection of data is the main problem of manual collection of error but it can be rectified successfully to involve the second one to the same project. Various type of error can be overcome if proper checking process can be initiate by appropriate body. For most critical data double check or triple check can be initiate. Growing continuous attention during data collection may reduce the data error. Sometime a review of the related data may decrease the data error. A color shading on projected excel sheet in the alternative row may help to track the mentioned data.

Cybersecurity Threats

Most of the time you can’t realize that a hacker or third party is trying to access your database. Person or individual who are trying to control your data send a short link with attractive or recent hot topics or similar to company email address link. Thousand of ways a hacker can try to damage or control your database system, so a strong security system shall be established to protect Data Integrity of the system.

Data Science Course

Failure to know the technical framework of database system, you can’t protect your data properly. If you are capable to handle the system then you need not to collect data science but if you want to update your existing your practice and eagerly want to update your team capability then you can involve your team in any data science course which are already available online which help your organization as well as development of self-confidence.

Devotion to Data Integrity

To keep subjects’ information safe & giving organization’s stakeholders the highest quality, accurate, complete, most data on which to base decisions need daily commitment for your Data integrity. A proper security system and group of trained individuals can support the organization in this situation to continue the company progress.

Data Integrity vs. Data Quality

Data quality is a crucial piece of the Data Integrity puzzle. It enables organizations to meet their data standards and ensure information aligns with their requirements with a variety of processes that measure data age, accuracy, completeness, relevance, and reliability. Data quality goes a step further by implementing processes and rules that govern data entry, storage, and transformation.

Data Integrity vs. Data Security vs. Data Quality

Data security involves protecting data from unauthorized access and preventing data from being corrupted or stolen. Data Integrity is typically a benefit of data security but only refers to data accuracy and validity rather than data protection.

Data Integrity & data security are more relevant to each other. Each of them plays a vital role for each other for their individual achievement. Data Integrity only refers to validation and accuracy of data but didn’t involve to protect data. On the other hand, data security confirms the protection against corruption or unauthorized access.

Data security plays a crucial role to maintain data integrity, on the other hand, Data Integrity is the end result of data security. To maintain data integrity, data security is the vital point and this situation may arise when accidental compromise occurs for data integrity.

Data Integrity is the essential component for the modern business procedure while making decision based on accuracy and efficiency of database. The main focus of the data security leads to Data Integrity and various type of procedure are applied to achieve the same.

Data quality can be defined that the data stored in database is compliant with the organization’s standards & requirements. It maintains integrity in a database. A set of rules to a specific or whole dataset and stores it in the target database shall be implement to do the same. Data Integrity shall be considered as data accuracy as well as correctness of data.

How to Protect Your data?

Validate Input

To ensure data accuracy data entry must be validated & verified input is significant when data is provided by familiar or unfamiliar sources, such as end-users, applications, & malicious users.

Remove Duplicate Data

Sensitive data stored in secure databases cannot be duplicated & it important to ensure that publicly available spreadsheets, emails, documents, & folders. To prevent unauthorized access to business-critical data or personally identifiable information duplicate shall be remove as soon as possible with the help of authorized personnel.

Back Up Data

To confirm data security & integrity data backups are crucial. To prevent the valuable data from permanent lost data backup shall be perform on regular manner at the end of everyday work. Organizations that suffer ransomware attacks, Data backups are especially important for them to protect their potential resources.

Access Controls

To maintaining data integrity, appropriate access controls shall be introduced in the organization. Data privileged option shall be implemented for specific user to control the database access procedure. This process will help the user understand their limitation to use the database system as well as to maintain the whole system.

Audit Trail

An audit trail is the standard practice to trace the unfavorable event. Data breach may occur at anytime in a renowned organization. If audit trail facility are available for the organization then it is very much to find out when and how data breached where. If proper information is available then it is easy to trac the source of attack. So audit shall be introduce at your organization for your database management system.

Assurance of Data Quality

It is the part of the Data Integrity process. Regular shall be conduct so that data can meet the certain standard. The processes of data accessibility, data cleaning, , data standardization is the main concern of the Data quality assurance. Data cleaning deal with inputting missing data, removing invalid entries, update same on timely manner.

Data accessibility deal with availability of the data to the stakeholders in secure and appropriate manner. For encoding and entering data, business shall be maintained and unauthorized data entry or transfer shall be prohibited. All type of company rule shall be implemented increase of transfer access of data to potential sector.

Data Corruption vs Data Integrity

Data corruption shall be considered as the serious Data Integrity failure. Based on the current practice, data corruption may be occurred through multiple channels. Most of them a very common problem is human error occur during collection or transferring of data. Malware and physical damage are another potential cause of data corruption.

Most human error often cause in wrong entry of collected data, unauthorized entry of database system, involving newcomer to sensitive practice with our prior training, programming etc.

It can be traced with appropriate data validation checks & restricting access to database system. Extensive & systematic use of backups can support restore databases in case of improper data entries.

Malware is another common cause of data corruption and this is basically occurred from external source which main purpose is to stole the data from potential data server. Cyberattacks are almost always unpredicted and instantly can’t recognized the source on the most of the cases. So here come the data encryption, always try to encrypt the critical and sensitive data and if possible tight security system shall be introduced though cost may be high to do the same.

To ensure organizational network security, regular penetration testing shall be done, this will help you to secure the organizational network system. A physical damage is may cause the data lose which mainly cause by accident and disasters. To protect data, data may be store in different physical location will protect from accident and natural disasters.

Data Integrity and Why Is It So Important? Read More »

Comparative Dissolution Study procedure

Comparative Dissolution, General Overview

[][]Comparative Dissolution, Oral dosage form like Table & Capsule are more popular than IV/IM(Intravenous/Intramuscular) Injection formulation. From the very beginning, people are most familiar with oral solid dosage form (Tablet, Capsule, Powder etc.) and becoming more popular till today as no special technique or device is not require to administer these products and associated pain is not involved here.

[][]In the period of time its are considered as the most effective and efficient method to treat the patient. This orally taken drugs are dissolved in GI (Gastro Intestinal) fluid and then bioavailable at the systemic circulation as it absorbed here. To measure the bioavailability of a certain drugs (in vivo Analysis, vivo is Latin for “within the living, test perform in living organism) is not accurately possible due to its complex nature.

[][]For this reason, in vitro (vitro is Latin for “within the glass, test perform outside the living organism) methods are followed to measure the dissolution rate of a certain drugs. This method is officially recognized by certain regulatory authority and it(in vitro study) considered most convenient way to develop new formulation of oral solid dosage form.

Comparative Dissolution Consideration

[][]Dissolution method is the best option for the lower strength drug where different strength is proportionally formulated to acquire the biowaiver of certain formulated drug. For a certain product which higher strength bioequivalence study has been carried out and found proportional to the concentration then biowaiver is conceivable to the lower strength.

Dissolution test are considering the most quality control tool for the commercial batch to batch product to monitor its consistency over a certain period of time. It also provides significant information during post approval changes of the certain product as changes made in formulation, manufacturing process and different scale up procedure.

[][]The most physiological factor is considered as the dissolution and solubility of the API and its permeability through the membrane of the GI[Gastro Intestinal] tract. As this measurement is so prone to error due to its complex nature then in vitro study consider the most convenient and reliable procedure to achieve the required target. During development of a certain solid dosage form, dissolution is considered as the best option to determine its quality parameter which have the great impact on the bioavailability of the formulated product.

Comparative Dissolution, Waiver of in vivo bioavailability

[][]BCS system applied in this case so that waiver for in vivo analysis can be assured. BCS [Biopharmaceutical Classification System] is a system which measure the permeability and solubility of drugs in a certain prescribed condition.

[][]The actual aim of BCS is to aid the post approval changes and arranging approval activities based on in vitro data studies.

This system has been optimized based on the oral solid dosage unit as most of the market products are available at oral dosage form [More than 50% total market share, US$23.4 Bn in 2021, US$24.7 Bn in 2022 as estimated, growth rate 5.9%).

[][]Waivers[ means giving permission to skip in vivo bioequivalence study] is actually reserved for those products that meet the specific requirements of solubility & permeability & most of the cases rapid dissolve in body fluid.

[][]Using the BCS, appropriate formulation study shall be developed such as Type II drugs designed as Permeable but insoluble, this class is not the actual right candidate for development of a new moiety.

[][]So, solubility shall be developed to acquire the right dissolution profile. Based on the solubility and permeability BCS has classified the four categories of the product as depicted below

Dosage form challenge

[][]Comparison has been drawn from old drug to new drug formulation, where older drugs compare to the current products are more prone to solubilities. Class II compound has been remarkably increased as 30% to 60% where class I compound has down to 40% to 20% where low solubility has the main cause to encounter the issue.

 A oral solid dosage form is the preferred option but all time this can’t possible the suspension or solution is continued to prove its existence  

[][]Generally a highly soluble active substance and rapidly dissolve dosage form provide better bioavailability and in this case biowaiver can be waived for bioequivalence studies base on its dissolution profile.

[][]If a active substance found low solubility but high permeability then the rate limiting steps of absorption may be consider as dissolution. Most of the cases dissolution profile control the more than one of excipients or special design matrix compounds. So Test condition may be consider as various time frame (10, 15, 20,30, 45 & 6 minutes).

[][]Drugs that are poor soluble in water then various time frame are considered and accepted timeframe is set for dissolution profile. Here USP Type 4 apparatus to be used to develop such type dissolution profile. Most of the time, monograph for combination product is not available at BP or USP the individual monograph shall be used to set the dissolution profile.

Selection of Dissolution Media

Selection of the dissolution media is the vital point to achieve the goal. pH of the media as the key role as all of the dosage form goes to GI[Gastro Intestinal] tract so pH shall be simulate with the GI Tract environment. pH shall be 1.2 to 6.8 which is the physiologic pH range of the body.

ZonepH
[A]Pre-prandial
Stomach
1.8(1~3)
Duodenum6.0(4~7)
Upper Jejunum 6.5(5.5~7)
Lower Jejunum 6.8(6~7.2)
Upper Ileum7.2(6.5~7.5)
Lower Ileum7.5(7~8)
Proximal Colon(5.5 ~6.5)
[B]Post-prandial
Stomach
4.0(3~6)
Duodenum5.0(4~7)
Upper Jejunum 5.5(5.5~7)
Lower Jejunum 6.5(6~7.2)
Upper Ileum7.2(6.5~7.5)
Lower Ileum7.5(7~8)
Proximal Colon(5.5 ~6.5)

Dissolution Statistics

Different cases obtaining after multipoint dissolution which is calculative as follows:
[][]If Test Product and Reference Product both shows dissolution rate more than 85% within first 15 minutes then no calculation is required, they are considered as similar. If it didn’t achieved then seek for next step.
[][]Seek for f2 value[ f2, similarity factor] if f2>50% then it consider similar then in vivo study is not required.
[][]Difference Factor [f1] is the percentage (%) difference between the two curve at the each time period and also measure the relative error between two curve.

How it works

To determine the difference and similarity factor(f2) following pont shall be noted:
[][]Use the Two different products for study, from each product collect 12 unit [12 unit from Test Product & 12 Unit from Reference Product].

[][]Three time point shall be considered[Exclude Point Zero], only one measurement after 85% shall be measured.
[][]Produced curve shall be similar, f2 values shall be close to 100. Most of the time f2 value more than 50 denote similarity of the two curves as well as equivalence of the two products.

 If Three/Four Time points come to the test then following points shall be considered. 

[][]The measurement for the Test Product and the Reference product shall be same. Dissolution Time point shall be same for the both product (10,15,20,30,45,60 minutes etc.). Products which tend to faster dissolution (85% dissolve within 30 minutes) then time frame shall be consider as 10, 15, 20, 30 minutes.
[][]Only One measurement shall be consider after completion of 85% dissolution of both sample and reference products.

System Requirements to Perform Comparative Dissolution

Dissolution Activities shall be continued on USP Type I Dissolution apparatus at 100 RPM or USP Type II Apparatus at 50 RPM using 900 ml of different dissolution media mentioned below.
[][]Media Use in Comparative Dissolution
[][]Acid Media: 0.1N HCL or Simulated Gastric Fluid USP without Enzyme
[][]Acetate Buffer pH 4.5
[][]Phosphate Buffer pH 6.8 or Simulated Intestinal Fluid without Enzyme
[][]If both the Test Product and Reference product shows more than 85% dissolution within first 15 minutes then no calculation required. If not meets the above requirements then calculate f2 Value.
[][]If found f2>50, then the profile considered as similar and in vivo study is not required. Minimum 12 unit of each shall be consider for comparative dissolution.

Comparative Dissolution Study procedure Read More »

Functions of Quality Assurance

Functions of Quality Assurance, Purpose:

Functions of Quality Assurance, The purpose of this SOP ( Standard Operating Procedure) is to outline the functions, principle duties and responsibilities of personnel working in Quality Assurance Department.

Functions of Quality Assurance, Scope:

To provide the guidelines for the proper implementation of the Quality Assurance System at XX Pharmaceuticals Limited ( Both General and Cephalosporin Block).

Definitions / Abbreviation:

[][]Quality Control Department : The Department carrying out day to day activities for the control of procedure, tests & analysis of products/ raw materials/ packaging materials/ stability samples and other relevant analysis of samples at laboratory to assist the manufacturing functions.
[][]Quality Assurance Department: The department carrying out day to day activities for the control of procedure, analysis of documents, regulatory functions, to assist the plant functions for the manufacturing, distributions and other relevant functions.
[][]CAPA : Corrective Action and Preventive Action.

Responsibilities:

[][]The roles and responsibility is as follows:

Executive, Quality Assurance

[][]To follow the guidelines of Quality Assurance
[][]To report their findings to their seniors

Asst. Manager, Quality Assurance

[][]To ensure implementation of the SOP.

AGM, Quality Assurance

[][]Approval of the SOP.

Procedure:

[][]Quality Assurance Department shall function for assuring the quality of all the manufactured batches and every stage of manufacturing / processing of drug Products.
[][]This shall be achieved by performing the functions of monitoring as per the laid down QA systems for the following areas:
[][]Monitor of incoming starting and packaging materials.
[][]Manufacturing process and process checks.
[][]Process monitoring and process control.
[][]Production Record Review.
[][]Final release or rejection of every batch of Drug Products for distribution and sale.
[][]Monitor stability testing and evaluation of shelf-life of products.
[][]Complaints handling and product recalls.
[][]Handling of Change Control Systems.
[][]Out of specification investigation.
[][]Investigation of Deviations.
[][]Returned products ( salvage and disposal ).
[][]Internal Quality Audits.
[][]Co-ordinate monthly Quality Review Meeting and implement the effective CAPAs regarding quality improvement.
[][]Control of non-confirming products.
[][]Co-ordinate all validation and qualification activities.
[][]Reprocessing of non-confirming products.
[][]To achieve the objectives of Quality Assurance functions of XX Pharmaceuticals Limited is classified as follows :
[][]To plan and manage all the activities of Quality Assurance Department to assure the quality of all products manufactured by the Company.
[][]To co-ordinate with manufacturing department in controlling their process and products at every stage of manufacturing to meet the established specifications though testing, auditing and reporting.
[][]To co-ordinate for the development of new product formulation, development of specifications, analytical procedure in co-ordination with Quality Control Department and Product Development Department.

[][]To review the adequacy and relevance of specifications & analytical procedures in co-ordination with Quality Control Department and Product Development Department.
[][]To co-ordinate technical audits of the Quality Control Laboratory to determine the analytical quality systems are yielding the highest quality information and to ensure that the analytical instrumentation is functioning properly and calibration and servicing is as per schedule.
[][]To be responsible for the release functions of the QA Department which shall include :
[][]Maintenance of Quality Control records of manufacturing procedures for each manufactured batch.
[][]Records of release, quarantine or rejection of components and finished products, containers, closures and labels based on Quality Control test results.

[][]Routine “Good Manufacturing Practices Auditing” of manufacturing process, control and related areas.
[][]To suggest and organize training program for the development of technical and administration skills of all the employees to meet with cGMP regulations on continuous basis, coordinating with GM, Plant and Quality Head.
[][]To assist Change Control Committee/Technical Committee for overall reviews of non-conformances, failure investigations, analyzing the Quality trends, investigations of market complaints, batch failure investigations, deviations, verifications of change control procedures, updating the specifications, test procedures, manufacturing processes etc.
[][]To liaison with regulatory authorities for new products manufacturing license and renewals of the same and other regulatory requirement.

Annexure:

N/A

Functions of Quality Assurance Read More »

Contract Manufacturing by Third Party

Contract Manufacturing, Purpose:

Contract Manufacturing, To establish a procedure that XX products are manufactured by the contractor using facilities and operations and to ensure that all batches of product manufactured at third party contract manufacturer that conforms to cGMP requirements undergo a formal evaluation of suitability for use prior to release to the market.

Contract Manufacturing, Scope:

The SOP is applicable for all products of XX Pharmaceuticals Limited manufactured & packed by a third party contract manufacturer.

Definitions / Abbreviation:

[][]QA – Quality Assurance
[][]QC – Quality Control
[][]QA – Quality Assurance
[][]BMR – Batch Manufacturing Record
[][]BPR – Batch Packaging Record
[][]ERP – Enterprise resource planning.
[][]cGMP – Current Good Manufacturing Practice
[][]COA – Certificate of Analysis

Responsibilities:

[][]The roles and responsibility is as follows:

Third party contractor

[][]To perform all the manufacturing related activities, analysis of the manufactured & packed product as per the contract/ toll manufacturing agreement

Executive, Production

[][]To check and review the production related activities.

Executive, Warehouse

[][]To supply and update the required raw and packaging materials as per forecast

Executive, QA

[][]To verify the overall manufacturing activities, collection of sample and related documents

General Manager, Plant

[][]Proper follow-up of overall activities

Manager, Quality Assurance

[][]Approval of all the toll manufacturing activities.

Procedure:

Selection of Third Party/ Contract Manufacturer

[][]Carry out GMP audit to assess the prospective contractor’s technical capability, capacity, cost effectiveness & the capability to maintain the required standards of cGMP & product quality.
[][]Document the business objectives and the justification for selecting a third party contractor.
[][]Review the manufacturing strategy which will be applied to the third party contractors.
[][]After developing a third party contract manufacturer, provide the product and their volume to the third party contractor for its technical assessment of manufacturing capability.
[][]Multiple strategies to be followed in selecting third party contractor.
[][]Consider the sources of materials which must meet the approved specification.
[][]Sign a contract manufacturing agreement with the selected contractor for manufacturing / packaging of products which will be followed accordingly.

[][]XX personnel will generate all the relevant documents as per contract manufacturer’s requirements for batch production.
[][]Production Executive will supply a production forecast of the required quantities of each product two months in advance so as to consider in third party’s

[][]production plan. XX shall have the right to revise the forecast in accordance with the market situation informing contractors one and half month in advance.
[][]Warehouse personnel of contract manufacturer will generate a month wise stock statement for raw and packaging materials to XX.
[][]Production Executive will generate the batch no. as per production forecast to Quality Assurance Executive along with requisition copy for BMR & BPR.

[][] QA Executive will issue the photocopy of BMR & BPR to Production Executive for manufacturing by third party. Master copy of all documents will be preserved at QA end.

[][]For testing procedure of raw and packaging materials will be followed as per contract manufacturing agreement. If any material (Raw/ Packaging) which will be tested by third party, will be forwarded by QC Executive from QUARANTINED stage along with forwarding letter to the contractor.
[][]After receiving the test report from third party, QC personnel will disposition the materials as PASSED/ REJECTED following the SOP.
[][]Production Executive along with warehouse Executive will supply all the materials to third party following the production forecast as per requirement within first week of running month. During supply of materials warehouse executive to be ensured the status label as per requirement by third party.
[][]Contract manufacturer will follow their production schedule and manufacturing operation will be started at the presence of Production and QA personnel of XX.

[][]XX personnel will verify all the written instruction in the BMR & BPR following the third party’s SOP as per cGMP requirement.
[][]All the in-process checks (IPC) will be conducted by contract manufacturer along with XX personnel.
[][]In case of any process validation activities will be conducted by XX personnel using the facility of third party.
[][]For testing of bulk intermediate and finished product, responsibility will be performed by XX and contract manufacturer as per contract manufacturing agreement.
[][]Sample for in-process check, chemical test and microbial test will be withdrawn as per agreement policy following the SOP of XX or contract manufacturer. [][]Finished product samples will be withdrawn as per same requirement.
[][]Retention sample will be kept at contract manufacturer’s end. If the retention sample is required for XX, then the sample will be forwarded to XX end by contract manufacturer following their own policy.
[][]Stability sample will be withdrawn from packaging line as per stability protocol and to be forwarded by contract manufacturer to XX end.

[][]If the stability test will be performed by contract manufacturer, samples will be kept in the manufacturer’s end as per protocol. In that cases test report will be forwarded to XX end accordingly. After receiving all the stability data, QA Executive will prepare the stability report.
[][]Third party contract manufacturer’s site will provide XX personnel with full batch documentation, copies of any unplanned event or deviation (including OOS), a copy of any investigation reports and COA reporting analytical test results against specification.
[][]QA and production personnel will review all the batch documents along with supportive records as per checklist for batch documentation after prior release from contract manufacturer.
[][]Original batch document or photocopy of the original will be provided by the contract manufacturer to XX end.

[][]Production personnel will issue the used quantity of raw and packaging materials in the ERP software with the help of IT personnel according to batch production record to update the material status.
[][]After reviewing the batch document, Production and QA representative will send the notification to the third party contract manufacturer about the batch disposition decision.
[][]Upon receiving the batch disposition decision from XX Pharmaceuticals Limited, the third party contract manufacturer will take step accordingly.
[][]XX shall take delivery of the finished products in XX’s own liabilities within fifteen days of intimation by contract manufacturer that the product is ready for dispatch.

[][]After completion of cost related process by both XX and contract manufacturer, central warehouse will receive the finished product and kept in the central warehouse of XX as per storage condition.

Annexure:

None

Contract Manufacturing by Third Party Read More »

Finished Product Handling, Storage & Distribution

Finished Product Handling, Purpose:

Finished Product Handling, The of these guidelines is to assist in ensuring the quality and identity of pharmaceutical products during all aspects of the distribution process such as storage, distribution, transportation, packaging, labeling, documentation and record-keeping practices.

Finished Product Handling, Scope:

This procedure is applicable for all finished products received and stored after receiving from Packaging department by Central Warehouse and different depots of XX Pharmaceuticals Ltd.

Definitions / Abbreviation:

[][]QA – Quality Assurance
[][]DIC – Depot In Charge
[][]ERP – Enterprise Resource Planning
[][]CWH – Central Ware House
[][]FIFO – First In First Out
[][]XX – Current version no. of the SOP

Responsibilities:

[][]The roles and responsibility is as follows:

Head of Distribution

[][]Head of Distribution shall responsible for overall the distribution activities.

Depot In Charge (DIC), Central Warehouse & Different Depots

[][]To verify, receive, storage and distribution the finished goods ensuring all the quality.

Head of Sales

[][]To monitor the distribution procedure as per market requirement.

Head of Marketing

[][]Prepare the market requirement for distribution and follow-up of overall activities.

General Manager, Plant

[][]Follow-up of overall activities.

Executive, Vat

[][]Follow the distribution activities as per Government legislation

Executive, QA

[][]Responsible for effective implementation and monitoring of procedure.
[][]Annual depot audit to verify the effectiveness of this procedure.

Manager, HR & Admin

[][]Shall responsible for monitor the effective distribution procedure

Manager, Quality Assurance

[][]To approve of all the distribution activities.

Procedure :

Precautions:

[][]The entire finished product should be handle carefully during distribution, storage, ensuring the quality and identity of the product.
[][]The entire finished product should be stored in product wise respective area maintaining product’s storage condition.
[][]During distribution FIFO should be controlled to avoid mix-up & errors.

Storage :

[][]Finished product will be received by central warehouse personnel from packaging department after getting the finished product release note from QA.
[][]Released finished product will be stored in the central warehouse at the product wise respective area as per storage condition.
[][]Warehouse personnel will monitor and record the temperature in “temperature monitoring record sheet ( Annexure – I)” for twice in a day.
[][]Storage areas should be designed or adapted to ensure appropriate and good storage conditions.
[][]In particular, they should be clean and dry and maintained within acceptable temperature limits. Products should be stored in the pallet which is suitably spaced to permit cleaning and inspection.
[][]Physical or other equivalent segregation should be provided for the storage of released, rejected, expired, recalled or returned products and suspected counterfeits.
[][]Broken or damaged items should be withdrawn from usable stock and stored separately up to final disposition.
[][]Storage areas should be provided with adequate lighting to enable all operations to be carried out accurately and safely.
[][]Periodic stock reconciliation should be performed by comparing the actual and recorded stocks. Stock discrepancies should be investigated by a multidisciplinary committee headed by HR & Admin.

Distribution

[][]All parties involved in the distribution of pharmaceutical products have a responsibility to ensure that the quality of pharmaceutical products and the integrity of the distribution chain are maintained throughout the distribution process from the site of the manufacturer to the entity responsible for dispensing or providing the product to the target consumer as per Government legislation.
[][]Head of Distribution send the depot wise market allocation to central ware house as per marketing/sales requirement.
[][]Central warehouse personal will prepare the released finished product as per market allocation for different depots along with necessary tertiary packaging.
[][]In-charge, CWH will prepare invoice/challan copy as well as ERP entry as per different depot requirement approved by head of Plant or his designee.
Executive, Vat will prepare the vat challan according to depot allocation ( through Musak-11) as per Government rule.
[][]After performing all the related activities, finished products will be transferred to different depot by company own transport or transport agent along with vat challan and invoice copy.
[][]DIC or designee will receive and check the finished pack according to invoice and to be stored following the procedure described on previous steps.

Market Dispatch :

[][]DIC will allocate the finished product as per market requirement for sale. The dispatch and transportation of products should be undertaken only after the receipt of a valid delivery order, which should be documented
[][]Dispatch record will be prepared by DIC and should include at least the following information:
[][]Date of dispatch;
=>Complete business name and address (no acronyms), type of entity responsible for the transportation, telephone number and names of contact persons;
=>Complete business name, address ( no acronyms), and status of the addressee (e.g. retail pharmacy, hospital or clinic);
=>A description of the products including, e.g. name, dosage form and strength ( if applicable);
=>Quantity of the products, e.g. number of containers and quantity per container ( if applicable);
=>Applicable transport and storage conditions;
=>A unique number to allow identification of the delivery order; and
=>Assigned batch number and expiry date ( where not possible at dispatch, this information should at least be kept at receipt to facilitate traceability).

Returned :

[][]If any products undergo as expired or packet damaged which is not fit for use then record the product and stored in a separate area of the depot/warehouse with proper labeling following annexure– II under custody of depot in-charge.
[][]DIC will make an invoice of those products and sent to central warehouse after entry in the ERP to adjust the stock. Returned products will be sent to CWH at every three months interval.
[][]Head of distribution will collect this invoice/challan from different depots and sent to central warehouse after getting prior approval from Head of Sales & Head of Marketing.

[][]After receiving the returned products from different depots, in-charge, CWH will report to QA department along with the approved invoice/challan copy. In-charge, CWH will segregate the returned products according to their nature in a separate area with proper labeling as per annexure-II.
[][]QA executive will take actions following the SOP ‘Disposition of Returned Finished Products.
[][]In case of market compliant or product recall, DIC will inform to Sales or Marketing Department following the respective SOPs Head of distribution will collect the market compliant sample (s) or recalled product(s), batch(s) within seven days of initiation.
[][]Head of QA will take actions for the market compliant and product recall procedure following SOP.
[][]Stock verification of different depots will make through a physical audit by the representatives of Audit & Accounts department once in a year.
[][]A quality audit will be conducted for all the depots headed by QA representative once in a year.

Annexure:

Annexure I – Temperature monitoring record sheet
Annexure II – Returned product label

Finished Product Handling, Storage & Distribution Read More »

Returned Finished Products Disposition

Returned Finished Products Disposition, Purpose :

Returned Finished Products Disposition, The objective is to specify a system to handle the returned finished products for their appropriate and adequate review and disposition in line with the Quality Management System.

Returned Finished Products Disposition, Scope :

This procedure is applicable for disposition of returned finished products and expired ones at XX Pharmaceuticals Limited. (Both General & Cephalosporin Block).

Definitions / Abbreviation:

[][]QA : Quality Assurance
[][]DIC : Depot In Charge
[][]XX : Current Version of SOP

Responsibilities:

[][]The roles and responsibility is as follows:

Head of Distribution

[][]Head of distribution shall collect the finished products from different depots and forward to the central warehouse for final disposition as per procedure.

Depot In Charge (DIC), Central Warehouse

[][]To verify, receive and separately quarantine the returned goods.

Executive, QA

[][]Responsible for effective implementation and monitoring of procedure.

Senior Executive, Cost & Budget (Factory)

[][]To verify all the returned products and make report for stock adjustment memo.

Head of Plant Operation

[][]Follow-up of overall activities.

Head of Quality Assurance

[][]Approval of all the disposition activities.

Procedure:

[][]The returned finished products shall either be destroyed or repacked and made fit for use depending upon the reason for which they have been returned. In either case, the associated risks must be well understood and addressed.
[][]Hence the procedure for disposition of returned finished products is defined depending upon the reason for which products are returned.
[][]Head of Distribution will collect the finished product as per requirement from different depots and send to Central warehouse following the Invoice/ Challan.
[][]On receipt of the products, warehouse (Central) In Charge or designee shall verify the products as per Invoice / Challan. Then warehouse in-charge will label the products to identify the primary cause of return and segregate the returned products in the respective area.
[][]Quarantine the products in area as per classification and inform QA and Head of Plant Operation through Returned Finished Product Note, (Annexure – I).
[][]Reference number for Returned finished products shall be alphanumeric system containing 7 characters. Numbering breakdown is as follows : RG/YY/001
For example : RG/XX/001
=>The first two alphabets shall stand for Returned Goods
=>Next two numeric characters shall stand for year code XX shall denote year 20XX.
=>Next character is followed by three numerical shall stand for serial number, which shall start every year
from 001.

[][]All the expired products returned from the market shall be directed and received at the distribution Department and these shall be destroyed in presence of QA Executive following the SOP of Procedure for Disposal of Materials & Products.
[][]QA Executive shall prepare Authorization and Disposal From for Materials & Products and take authorization for disposal activities.
[][]Head of Quality Assurance or designee shall arrange to verify the reason for return/ recall of the returned products against the intimation.
[][]If the returned products (except expired products) which are fit for further use they shall be repacked through repackaging request form following the Reprocessing/reworking and Repackaging Procedure.
[][]Quality Assurance personal shall examine the products returned because of dirty labeling, packed damaged products can be repacked. To ensure the product quality, QA executive shall draw the sufficient sample for complete analysis (If necessary) and only if found satisfactory and certified accordingly, then the batch shall be allowed for repacking.
[][]For the repackaging activities minimum five unit packs are required for the returned product of same batch no. Less than five unit pack of a same batch shall be disposed following the disposal SOP.
[][]The returned products except expired products which can be repacked to be sent from central warehouse to the manufacturing site and shall be classified and stored separately depending upon the primary cause of return.
[][]Record for issue of packaging material, overprinting and packing shall be maintained on Additional Batch Packaging Record (Primary/Secondary) along with the Repackaging Request Form authorization by Head of Quality Assurance.
[][]If the returned products in the opinion of the Quality Assurance personnel are not considered fit for repackaging, the same shall be destroyed after approval from Head of Quality Assurance.
[][]This repacked part shall be released after compiling of the all related documents and shall be attached to the earlier batch record (BMR/BPR)
Products recalled voluntarily or recalled, as per the directives from Drug Control Authorities the recalled products shall be handled depending upon the reason for product recall; they may be require to be destroyed or repacked.

Annexure:

Annexure I – Returned Finished Product Note

Returned Finished Products Disposition Read More »

Cleaning Validation Procedure

Cleaning Validation, Purpose:

Cleaning Validation, To lay down a procedure for carrying out cleaning validation to establish validated cleaning procedure for manufacturing and primary packaging equipment and effectively maintain cleaning process in validated state. Cleaning must be demonstrated to be effective in order to provide assurance that unacceptable levels of contamination are not carried over into subsequent products. It defines the cleaning validation and verification requirements to ensure that all cleaning procedures which may impact product quality are formally validated or verified. Cleaning must be demonstrated to be effective in order to provide assurance that unacceptable levels of contamination are not carried over into subsequent products. It defines the cleaning validation and verification requirements to ensure that all cleaning procedures which may impact product quality are formally validated or verified.

Cleaning Validation, Scope:

This procedure is applicable for equipment’s used in product manufacturing and primary packaging of XX Pharmaceuticals Limited (Both General & Cephalosporin Block).

Definitions / Abbreviation:

[][]QA : Quality Assurance
[][]QC : Quality Control
[][]CIP : Clean-in-place
[][]OOS : Out of Specification
[][]MACO : Maximum Allowable Carryover

Responsibilities:

[][]The roles and responsibility is as follows:

Concerned Department

[][]To cooperate the validation team to make the cleaning validation job successful.

Quality Assurance

[][]Responsible for the developing and co-ordinate the entire cleaning validation activities as a team leader.

Quality Control

[][]Ensure that analytical method has been prepared and validated before conducting cleaning validation and coordinate all analytical & Microbiological test.
To coordinate all the sampling activities.

Head of Plant Operation

[][]Proper follow-up of overall activities.

Head of Quality Assurance

[][]Approval of all the validation activities.

Procedure:

[][]Prior to conducting a cleaning validation study on an Active Ingredient (of previous product) following activities need to be summarized. A Validation team shall be formed comprising of members from Production, Engineering, Quality Control, Validation / Quality Assurance.

Defining the key elements

[][]Define solubility, toxicity of active ingredient in cleaning validation protocol. If any of the active ingredients are deemed to be more potent or less soluble in water than the ingredients for which cleaning validation has already been completed, or deemed to be particularly difficult to clean from equipment, a new validation study will be carried out.
[][]Define the product-equipment matrix.
[][]Prepare the equipment matrix. Determine the total contact surface area of the equipment/ instruments which are to be used in the manufacturing of that product with the help of Engineering and Production Department.
[][]Determine the scope and establish acceptance criteria.
[][]Develop and validate sampling procedure and analytical method.
[][]Designing the cleaning validation protocol. All protocols shall be as per the Format given I Annexure- I.
[][]Execute the cleaning validation study and protocol.
[][]Summarize the data and compilation of the final report.
[][]Devising and monitoring program to establish that the cleaning process is continued to be in validated state.
[][]Establishing conditions for which re-validation would be required.

Describing the key elements

[][]The product – equipment matrix
[][]Prepare a Product–equipment matrix (Annexure-II) & Equipment matrix (Annexure-III) for the products, equipments used in the cleaning validation study.
[][]The matrixes will provide information about the manufacturing and packing line used for multiple products and possible product contact surface area.
[][]The matrix will indicate the worst case at a glance and justify the acceptance criteria for a cleaning validation.

Determining the scope

[][]The scope includes evaluation of residual contamination of active ingredient.
[][]Cleaning validation to be done based on matrix for including any new product in to the matrix for supplying product to local market. However, all products deemed to be supplied to highly regulated markets, the cleaning validation to be done for every product.
[][]Acceptance criteria for Residual active ingredients: The approach for setting limits can be (1) product specific cleaning validation for all products; (2) grouping into product families and choosing a worst case product; (3) grouping into groups of risk (e.g. cleaning difficulty, solubility, toxicity potency of API, facility volume of product); (4) setting limits on not allowing more than a certain fraction of carryover; (5) different safety factors for different dosage forms.
[][]For determining cleaning validation process, widely used criteria, i.e. 1/1000, will be consider for MACO calculation and to evaluate the cleaning effectiveness.
Establish the limit for Maximum Allowable Carryover (MACO) according to the following equation.

MACO = TDD Previous X MBS/ SF X TDD Next

Where,
=>MACO = Maximum Allowable Carryover: acceptable transferred amount from the investigated product ( “previous” )
=>TDD Previous = Standard therapeutic dose of the investigated product ( in the same dosage form as TDD next)
=>TDD Next = Standard therapeutic dose of the daily dose for the next product.
=>MBS = Minimum Batch Size for the next product(s) (where MACO can end up)
=>SF = Safety Factor ( normally 1000 is used in calculation based on TDD )
#(Ref. APIC : Active Pharmaceutical Ingredients Committee)

[][]However, the other criteria i.e. 10 ppm Criteria, API toxicity data will also be taken into consideration for evaluation of cleaning effectiveness. The cleaned equipment’s will also be checked visually to ensure the cleaning effectiveness.
[][]1/1000 Criteria: NMT 0.1% of the normal therapeutic dose of any product to appear in the maximum daily dose of the following product.
[][]10 ppm Criteria : NMT 10 ppm of any product to appear in another product;
[][]Based on toxicity : LD50 Value of API will be considered;
[][]Visual Inspection: No quantity of residue to be visible on the equipment after cleaning procedures is performed.
[][]API Solubility: The solubility of drug in washing solvent plays a great role in cleaning. The lesser the solubility greater will be the difficulty to remove the residue from surface.
[][]Thus least soluble molecule (based on the pharmacopoeia or other reliable reference) will be taken for the analysis since if least soluble molecule gives satisfactory result in cleaning validation and routine monitoring then we can rest assure for the other higher soluble drug.
[][]API Toxicity: An evaluation according to the toxicities should be carried out based on the material safety data sheet (MSDS).
[][]Other risk factors such as potency and facility volume of a particular product should also be taken into consideration during designing cleaning validation program.
[][]Develop and validate analytical method for cleaning validation sample analysis.

Swab sampling

[][]Generally method involves swabbing of 5 cm2 but larger area could be considered as per regulatory dossier or with proper justification and difficult to clean areas will be considered as per below Diagrammatic representation.
[][]Define the most difficult to clean areas in the piece of equipment (show delineate pictorially as far as possible).
[][]A polyester tipped swab (Texwipe, Alpha swab with long handle) shall be used. One side of swab shall be given horizontal 10 strokes and then reverse side of the swab shall be given vertical 10 strokes.
[][]The swab stick shall be dipped into the screw cap test tubes/ vials containing solvent (Approximately 10 ml). One test tube / vial will be taken for each swab sample.
[][]The test tube / vial will be labeled specifying the particular piece of equipment swabbed & the location (where appropriate).
[][]The soaked swab (in solvent) to be firmly squeezed along the side of the test tubes / vials and then sampling side to be swabbed. After taking samples from the equipment/utensils the swab to be returned to the respective screw cap test tube/vial.

[][]Each vial will be closed with swab inside & sent to QC for analysis of the swab sample.
[][]Study and establish the stability of sample solution storage condition & storage time, based on a protocol.
[][]Perform the recovery analysis on the swab with spiked SS surface/ similar to the equipment material of construction. Recovery of drug shall be not less than 85% after taking into account interference of swabbing material.
[][]Perform the recovery study of the API from the swab stick for record only.
[][]Mention the swab sampling point details in the cleaning validation protocol.

Rinse sampling

[][]For rinse sample collection graduated plastic mug/container, plastic squeezer, stainless steel container to be used, will be cleaned first with 70% IPA followed by sufficient purified water (Wipe with lint free cloth soaked in 70% IPA if required after visual inspection).
[][] After collection of each rinse the required quantity will be transferred to the conical flask for QC submission. To discard the leftover residue in the SS container/ plastic mug these utensils will be cleaned with sufficient purified water before next use.
[][]Same procedure will be followed for MDI unit except for cleaning purpose only absolute ethanol will be used instead of 70% IPA and purified water.
[][]Rinse sample can be evaluated at intervals during the cleaning and at the completion of the cleaning process. Collection of rinse samples should consider location, timing and volume.
[][]Rinse sample allow sampling of a large surface area and of inaccessible systems or ones that cannot be routinely disassembled. However consideration should be given to the fact that the residue or contaminant may be insoluble or may be physically occluded in the equipment.
[][]Study and establish the stability of sample storage conditions and storage time, based on a protocol.
[][]For those equipment or parts (e.g. turret, die and punch of compression machine, hopper) rinse sampling method is not feasible, only swab sampling method will be applied.
[][]At first swab sample will be collected as per plan and will be sent to QC. Then the swab taken area will be cleaned with purified water and finally with lint free cloth.
[][]At first the equipment will be cleaned as per the equipment cleaning SOP and the dismantled parts will be stored in the clean equipment store room after drying.
[][]After collecting rinse samples the cleaned equipment’s will be allowed for self drying before next use. Finally the rinse sample in a closed conical flask with proper labeling will be sent to QC for analysis.

[][]Perform the recovery analysis on the rinse with spiked SS surface/ similar to the equipment material of construction. Recovery of drug shall be not less than 85%.
[][]Microbiological considerations: weather or not CIP systems are used for cleaning of processing equipment, consider the microbiological aspects of equipment cleaning.
[][]The equipment/utensils are considered clean if the total microbial count is less then 25 CFU per 25 cm2/ per swab or 100 cfu/ml taken from equipment/utensils surface and there are no objectionable organisms present ( e.g. E.coli, Salmonella, Pseudomonas aeruginosa, Staphylococcus aureus).

[][]If any growth is observed, appropriate tests to identify the organisms are to be conducted. If the acceptance limit is exceeded, or an objectionable microorganism is identified, then then an OOS investigation must be initiated by the Microbiological Laboratory to identify the source of microbial contamination. Investigation should be done by the manufacturing department.

[][]Contaminated equipment must be re-cleaned and re-swabbed/re-rinsed and it be must meet acceptance limits prior to release for further manufacturing.
For all the types of equipment initial cleaning validation shall be performed on at least three consecutive batches or three runs.

Bracket Approach

[][]The design uses the extremes to cover in between range. That is the largest pieces of identical equipment could cover all of the in between sizes.
[][]The lowest and highest strength of the products.

Worst Case Approach

[][]The most difficult to clean equipment could cover all the easier to clean equipment also.

[][]Initially cover all the types of equipment for three runs.
[][]Type A cleaning done after 6 days (144 hours) from the first day of use of the equipment or area of solid dosage forms.
[][]The dirty holding period of manufacturing equipment should not be more than 5 days.
[][]In case of batch size change, either increase or reduction in the batch size, the matrix shall be re-evaluated.
[][]If a drug product is manufactured in different strengths, cleaning procedures shall be validated for the highest strength of that particular drug product. If the manufacturing line contains multiple products or a single product contains multiple active ingredients, the active ingredient which is least soluble/more in concentration and/or more toxic, hard to clean shall be taken into consideration for cleaning validation/verification.
[][]If there are multiples of particular equipment with same configuration, material of construction and cleaning procedure then cleaning validation of one piece of that particular equipment will be adequate.
[][]If there are different sizes of the same equipment with same cleaning procedure, cleaning validation for the largest size of that particular equipment is adequate.
[][]The worst case selection criteria will be a) Hard to clean product (Color and Flavor) b) Least soluble API c) Maximum toxicity of API d) Minimum therapeutic daily dose e) Facility volume of a particular product.

Execution of cleaning validation protocol/study

[][]Adequate training to the operators (Production, QC and Engineering) shall be given and documented to execute the protocol efficiency.
[][]Ensure that during validation study the results and activities represent regular daily operations.
[][]Ensure that the same piece of equipment is available for the execution of the cleaning by the operators.
[][]Ensure that operator’s variability is addressed during the process design to maximize the representation of results.

=>Cleaning Validation Protocol number shall be given as follow;
CVR/XXX/YY
Where
=>CVP stands for Cleaning Validation Protocol/stands for separator
=>XXX stands for the sequential number which starts from 001 for calendar year.
=>YY stands for the last two digit of the year.

Summarize the data and generate the report

[][]The final result of particular sampling site shall be ‘difficult to clean’ reported and incorporated in the calculations.
[][]Test results shall be complied by Validation/QA.
[][]Any change in procedure shall be approved by QA.
[][]Evaluation of product matrix shall be done periodically or during introduction of any new product into the existing facility.
[][]Objective & Scope: Describe the objective and scope of the cleaning validation / verification in cleaning validation /verification protocol.
[][]Cleaning Validation/Verification Protocol: provides the tabulated test results and the acceptance criteria for each piece of equipment and equipment train.
[][]Cleaning Validation/Verification process shall be concluded with a summarized report incorporated as a final report, which is written and approved.
[][]Tabulation of test results: provides the tabulated test results and mention the acceptance criteria for each piece of equipment and equipment train.
[][]Discussion and Conclusion: discuss the cleaning validation program under study and summarize the outcome of the cleaning validation program.
[][]Cleaning Validation Report number will be same as protocol no. given as follows.
=>CVR/XXX/YY
=>Where, CVR stands for Cleaning Validation Report.
=>Revalidation of cleaning procedure is required in case of following change
=>Equipment or equipment configuration changes.
=>Changes in cleaning procedure and change of formulation where active quality is increased (e.g. overage).
=>Change in acceptance criteria due to change in batch size or introduction of new product (the matrix shall be Re-evaluated by preparing a separate cleaning validation protocol).
=>Process change in manufacturing process (e.g. wet granulation to direct compression and vice versa).
=>The significance of change shall be evaluated collectively by Production, Engineering and QA.
=>Whenever any change control is made to a validated cleaning procedure, due to change in equipment, process, repair or any other valid reason, then the need evaluation shall be done jointly by the Head of Production, Head of Engineering and finally approved by Manager QA.
=> If the decision is made to revalidate, the same be implemented.

Annexure:

Annexure I-Cleaning Validation Protocol
Annexure II-Product Equipment Matrix
Annexure III-Equipment Matrix

Cleaning Validation Procedure Read More »